return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H7NO (Oxazole, 4,5-dihydro-2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-283.839980
Energy at 298.15K-283.848883
HF Energy-283.246911
Nuclear repulsion energy235.689879
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3208 3051 4.03      
2 A 3135 2982 24.08      
3 A 3126 2974 25.64      
4 A 3106 2955 5.05      
5 A 1657 1576 39.18      
6 A 1613 1534 0.14      
7 A 1589 1511 2.65      
8 A 1575 1498 18.35      
9 A 1487 1414 12.94      
10 A 1379 1312 9.23      
11 A 1341 1276 2.49      
12 A 1240 1180 97.08      
13 A 1090 1037 8.96      
14 A 950 904 43.42      
15 A 914 870 3.28      
16 A 877 835 46.03      
17 A 847 806 14.70      
18 A 734 698 9.92      
19 A 644 612 4.01      
20 A 333 317 5.04      
21 A 3199 3043 34.46      
22 A 3175 3020 0.11      
23 A 3172 3018 6.15      
24 A 1581 1504 8.97      
25 A 1297 1233 0.20      
26 A 1217 1157 0.21      
27 A 1141 1085 2.78      
28 A 1110 1056 0.73      
29 A 855 813 0.13      
30 A 567 540 4.76      
31 A 237 226 13.07      
32 A 142 135 1.40      
33 A 120 114 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 24328.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23143.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.24678 0.10956 0.07932

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.372 2.159 0.000
C2 0.528 -1.557 0.000
C3 -1.025 -1.316 0.000
N4 -1.191 0.208 0.000
O5 1.120 -0.171 0.000
C6 0.000 0.706 0.000
H7 0.875 -2.071 0.898
H8 0.875 -2.071 -0.898
H9 -1.508 -1.718 0.893
H10 -1.508 -1.718 -0.893
H11 0.969 2.390 0.887
H12 0.969 2.390 -0.887
H13 -0.543 2.753 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 N4 O5 C6 H7 H8 H9 H10 H11 H12 H13
C13.71993.74552.50012.44771.50004.35354.35354.40014.40011.09411.09411.0910
C23.71991.57232.46471.50732.32431.09101.09102.22892.22894.06994.06994.4416
C33.74551.57231.53312.43202.26712.23332.23331.09171.09174.30104.30104.0974
N42.50012.46471.53312.34281.29133.20483.20482.14642.14643.19613.19612.6261
O52.44771.50732.43202.34281.42302.11542.11543.17733.17732.71492.71493.3644
C61.50002.32432.26711.29131.42303.04683.04682.99092.99092.13592.13592.1179
H74.35351.09102.23333.20482.11543.04681.79612.40853.00164.46204.80595.1076
H84.35351.09102.23333.20482.11543.04681.79613.00162.40854.80594.46205.1076
H94.40012.22891.09172.14643.17732.99092.40853.00161.78664.79665.11644.6600
H104.40012.22891.09172.14643.17732.99093.00162.40851.78665.11644.79664.6600
H111.09414.06994.30103.19612.71492.13594.46204.80594.79665.11641.77451.7905
H121.09414.06994.30103.19612.71492.13594.80594.46205.11644.79661.77451.7905
H131.09104.44164.09742.62613.36442.11795.10765.10764.66004.66001.79051.7905

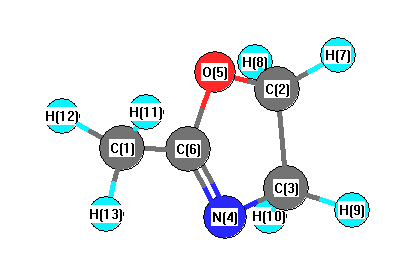
picture of Oxazole, 4,5-dihydro-2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C6 N4 127.032 C1 C6 O5 113.709
C2 C3 N4 105.056 C2 C3 H9 112.322
C2 C3 H10 112.322 C2 O5 C6 104.933
C3 C2 O5 104.293 C3 C2 H7 112.725
C3 C2 H8 112.725 C3 N4 C6 106.459
N4 C3 H9 108.548 N4 C3 H10 108.548
N4 C6 O5 119.259 O5 C2 H7 107.936
O5 C2 H8 107.936 C6 C1 H11 109.860
C6 C1 H12 109.860 C6 C1 H13 108.621
H7 C2 H8 110.799 H9 C3 H10 109.815
H11 C1 H12 108.384 H11 C1 H13 110.055
H12 C1 H13 110.055
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability