return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

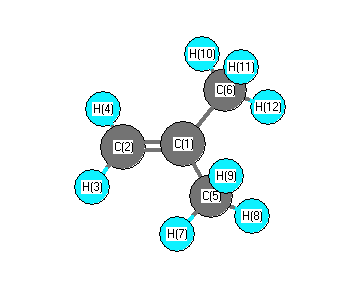
All results from a given calculation for CH2C(CH3)CH3 (1-Propene, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-155.610536
Energy at 298.15K-155.618461
HF Energy-155.245341
Nuclear repulsion energy118.747903
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3165 3011 9.78      
2 A1 3154 3000 18.76      
3 A1 3071 2921 17.87      
4 A1 1716 1632 2.83      
5 A1 1601 1523 9.76      
6 A1 1523 1449 0.18      
7 A1 1496 1423 0.99      
8 A1 1134 1078 4.40      
9 A1 790 752 0.00      
10 A1 383 365 0.23      
11 A2 3128 2976 0.00      
12 A2 1577 1500 0.00      
13 A2 1086 1033 0.00      
14 A2 726 691 0.00      
15 A2 180 171 0.00      
16 B1 3131 2979 40.61      
17 B1 1592 1515 16.76      
18 B1 1170 1113 0.99      
19 B1 932 887 50.12      
20 B1 429 408 7.63      
21 B1 198 189 0.63      
22 B2 3248 3090 18.06      
23 B2 3152 2998 17.04      
24 B2 3068 2918 16.96      
25 B2 1594 1516 4.07      
26 B2 1482 1410 9.64      
27 B2 1318 1254 5.99      
28 B2 1032 982 0.38      
29 B2 980 932 0.20      
30 B2 449 427 0.80      

Unscaled Zero Point Vibrational Energy (zpe) 24251.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23070.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.29943 0.27525 0.15159

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.127
C2 0.000 0.000 1.471
H3 0.000 0.924 2.043
H4 0.000 -0.924 2.043
C5 0.000 1.287 -0.689
C6 0.000 -1.287 -0.689
H7 0.000 2.165 -0.034
H8 0.887 1.330 -1.334
H9 -0.887 1.330 -1.334
H10 0.000 -2.165 -0.034
H11 -0.887 -1.330 -1.334
H12 0.887 -1.330 -1.334

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 H7 H8 H9 H10 H11 H12
C11.34372.12702.12701.52451.52452.17122.16632.16632.17122.16632.1663
C21.34371.08701.08702.51482.51482.63703.22893.22892.63703.22893.2289
H32.12701.08701.84892.75633.51522.41963.51523.51523.72304.15654.1565
H42.12701.08701.84893.51522.75633.72304.15654.15652.41963.51523.5152
C51.52452.51482.75633.51522.57451.09541.09741.09743.51402.83802.8380
C61.52452.51483.51522.75632.57453.51402.83802.83801.09541.09741.0974
H72.17122.63702.41963.72301.09543.51401.78131.78134.33033.83343.8334
H82.16633.22893.51524.15651.09742.83801.78131.77363.83343.19772.6608
H92.16633.22893.51524.15651.09742.83801.78131.77363.83342.66083.1977
H102.17122.63703.72302.41963.51401.09544.33033.83343.83341.78131.7813
H112.16633.22894.15653.51522.83801.09743.83343.19772.66081.78131.7736
H122.16633.22894.15653.51522.83801.09743.83342.66083.19771.78131.7736

picture of 1-Propene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H3 121.745 C1 C2 H4 121.745
C1 C5 H7 110.877 C1 C5 H8 110.370
C1 C5 H9 110.370 C1 C6 H10 110.877
C1 C6 H11 110.370 C1 C6 H12 110.370
C2 C1 C5 122.394 C2 C1 C6 122.394
H3 C2 H4 116.511 C5 C1 C6 115.212
H7 C5 H8 108.659 H7 C5 H9 108.659
H8 C5 H9 107.823 H10 C6 H11 108.659
H10 C6 H12 108.659 H11 C6 H12 107.823
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability