return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10S2 (1,4-Butanedithiol)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-948.367280
Energy at 298.15K-948.378057
HF Energy-947.756896
Nuclear repulsion energy332.912368
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3107 2956 0.00      
2 Ag 3082 2932 0.00      
3 Ag 2781 2646 0.00      
4 Ag 1586 1509 0.00      
5 Ag 1583 1506 0.00      
6 Ag 1435 1365 0.00      
7 Ag 1357 1291 0.00      
8 Ag 1108 1054 0.00      
9 Ag 1046 995 0.00      
10 Ag 875 833 0.00      
11 Ag 773 736 0.00      
12 Ag 343 326 0.00      
13 Ag 214 203 0.00      
14 Au 3170 3016 37.76      
15 Au 3138 2986 4.52      
16 Au 1392 1324 1.56      
17 Au 1183 1125 2.89      
18 Au 948 902 2.24      
19 Au 772 735 5.28      
20 Au 193 183 39.36      
21 Au 107 102 17.64      
22 Au 57 54 5.80      
23 Bg 3164 3010 0.00      
24 Bg 3121 2969 0.00      
25 Bg 1399 1331 0.00      
26 Bg 1331 1266 0.00      
27 Bg 1109 1055 0.00      
28 Bg 825 784 0.00      
29 Bg 186 177 0.00      
30 Bg 139 133 0.00      
31 Bu 3109 2957 45.69      
32 Bu 3089 2938 7.06      
33 Bu 2781 2646 21.80      
34 Bu 1599 1522 6.38      
35 Bu 1579 1502 8.33      
36 Bu 1413 1344 40.52      
37 Bu 1307 1243 20.70      
38 Bu 1045 995 3.71      
39 Bu 905 861 2.53      
40 Bu 738 702 2.85      
41 Bu 405 385 6.27      
42 Bu 105 99 3.79      

Unscaled Zero Point Vibrational Energy (zpe) 29799.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 28348.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.47091 0.01850 0.01804

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.372 3.172 0.000
S2 -1.372 -3.172 0.000
C3 1.444 1.333 0.000
C4 -1.444 -1.333 0.000
C5 0.000 0.775 0.000
C6 0.000 -0.775 0.000
H7 2.694 3.382 0.000
H8 -2.694 -3.382 0.000
H9 -0.530 1.142 0.887
H10 -0.530 1.142 -0.887
H11 0.530 -1.142 0.887
H12 0.530 -1.142 -0.887
H13 -1.972 -0.984 -0.892
H14 -1.972 -0.984 0.892
H15 1.972 0.984 -0.892
H16 1.972 0.984 0.892

Atom - Atom Distances (Å)
  S1 S2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S16.91111.84005.31222.76184.17781.33887.71252.92012.92014.48324.48325.40795.40792.43772.4377
S26.91115.31221.84004.17782.76187.71251.33884.48324.48322.92012.92012.43772.43775.40795.4079
C31.84005.31223.93031.54822.55482.40046.27342.17322.17322.78312.78314.22294.22291.09381.0938
C45.31221.84003.93032.55481.54826.27342.40042.78312.78312.17322.17321.09381.09384.22294.2229
C52.76184.17781.54822.55481.54923.74934.95341.09711.09712.17742.17742.78872.78872.17462.1746
C64.17782.76182.55481.54821.54924.95343.74932.17742.17741.09711.09712.17462.17462.78872.7887
H71.33887.71252.40046.27343.74934.95348.64794.02554.02555.09245.09246.45226.45222.65872.6587
H87.71251.33886.27342.40044.95343.74938.64795.09245.09244.02554.02552.65872.65876.45226.4522
H92.92014.48322.17322.78311.09712.17744.02555.09241.77472.51783.08043.12442.56833.07482.5077
H102.92014.48322.17322.78311.09712.17744.02555.09241.77473.08042.51782.56833.12442.50773.0748
H114.48322.92012.78312.17322.17741.09715.09244.02552.51783.08041.77473.07482.50773.12442.5683
H124.48322.92012.78312.17322.17741.09715.09244.02553.08042.51781.77472.50773.07482.56833.1244
H135.40792.43774.22291.09382.78872.17466.45222.65873.12442.56833.07482.50771.78384.40794.7552
H145.40792.43774.22291.09382.78872.17466.45222.65872.56833.12442.50773.07481.78384.75524.4079
H152.43775.40791.09384.22292.17462.78872.65876.45223.07482.50773.12442.56834.40794.75521.7838
H162.43775.40791.09384.22292.17462.78872.65876.45222.50773.07482.56833.12444.75524.40791.7838

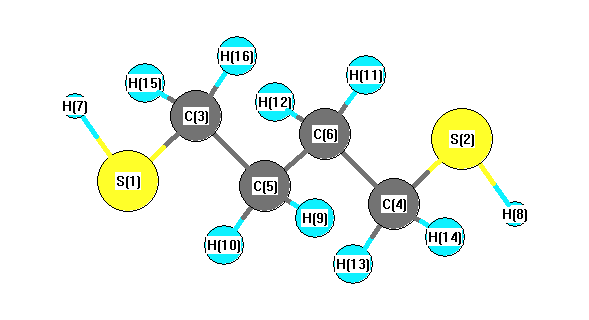
picture of 1,4-Butanedithiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 108.893 S1 C3 H15 109.747
S1 C3 H16 109.747 S2 C4 C6 108.893
S2 C4 H13 109.747 S2 C4 H14 109.747
C3 S1 H7 96.804 C3 C5 C6 111.136
C3 C5 H9 109.290 C3 C5 H10 109.290
C4 S2 H8 96.804 C4 C6 C5 111.136
C4 C6 H11 109.290 C4 C6 H12 109.290
C5 C3 H15 109.591 C5 C3 H16 109.591
C5 C6 H11 109.548 C5 C6 H12 109.548
C6 C4 H13 109.591 C6 C4 H14 109.591
C6 C5 H9 109.548 C6 C5 H10 109.548
H9 C5 H10 107.969 H11 C6 H12 107.969
H13 C4 H14 109.258 H15 C3 H16 109.258
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability