return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-289.429010
Energy at 298.15K-289.446211
HF Energy-288.763860
Nuclear repulsion energy332.819306
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3181 3026 38.50      
2 A 3159 3005 2.85      
3 A 3133 2980 2.00      
4 A 3091 2941 2.59      
5 A 2993 2847 144.72      
6 A 1612 1534 6.99      
7 A 1594 1517 2.46      
8 A 1587 1510 0.30      
9 A 1487 1415 5.64      
10 A 1463 1391 10.68      
11 A 1368 1302 1.19      
12 A 1220 1160 7.30      
13 A 1129 1074 3.74      
14 A 1007 958 1.19      
15 A 853 811 1.18      
16 A 711 676 11.38      
17 A 429 408 4.82      
18 A 314 299 1.54      
19 A 190 181 0.93      
20 A 88 84 0.02      
21 E 3183 3028 21.69      
22 E 3163 3009 44.68      
23 E 3131 2978 19.09      
24 E 3091 2941 19.95      
25 E 2988 2843 9.26      
25 E 2988 2843 9.26      
26 E 1610 1531 3.53      
26 E 1610 1531 3.53      
27 E 1600 1522 2.99      
27 E 1600 1522 2.99      
28 E 1587 1510 1.77      
28 E 1587 1510 1.77      
29 E 1486 1413 7.54      
29 E 1486 1413 7.54      
30 E 1438 1368 10.26      
31 E 1372 1305 16.85      
32 E 1265 1203 13.82      
33 E 1142 1087 3.69      
33 E 1142 1087 3.69      
34 E 1072 1020 31.19      
35 E 933 888 0.67      
35 E 933 888 0.67      
36 E 840 799 4.41      
36 E 840 799 4.41      
37 E 473 450 3.95      
37 E 473 450 3.95      
38 E 307 292 0.98      
38 E 307 292 0.98      
39 E 230 219 0.01      
39 E 230 219 0.01      
40 E 89 85 0.34      
40 E 89 85 0.34      

Unscaled Zero Point Vibrational Energy (zpe) 46302.2 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 44047.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.07561 0.07561 0.04326

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.018
C2 0.354 1.384 -0.448
C3 1.022 -0.998 -0.448
C4 -1.375 -0.386 -0.448
C5 -0.333 2.453 0.428
C6 2.291 -0.938 0.428
C7 -1.958 -1.515 0.428
H8 1.438 1.496 -0.348
H9 0.103 1.532 -1.515
H10 0.576 -1.993 -0.348
H11 1.275 -0.855 -1.515
H12 -2.014 0.497 -0.348
H13 -1.378 -0.676 -1.515
H14 -1.422 2.413 0.329
H15 -0.071 2.271 1.474
H16 0.000 3.456 0.138
H17 2.801 0.025 0.329
H18 2.003 -1.074 1.474
H19 2.993 -1.728 0.138
H20 -1.378 -2.438 0.329
H21 -1.931 -1.197 1.474
H22 -2.993 -1.728 0.138

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.50261.50261.50262.50962.50962.50962.10702.16962.10702.16962.10702.16962.81792.69873.45832.81792.69873.45832.81792.69873.4583
C21.50262.47412.47411.54403.14863.81031.09451.10573.38592.64572.53062.89532.19462.15932.18242.90463.52904.12244.26753.94714.6076
C31.50262.47412.47413.81031.54403.14862.53062.89531.09451.10573.38592.64574.26753.94714.60762.19462.15932.18242.90463.52904.1224
C41.50262.47412.47413.14863.81031.54403.38592.64572.53062.89531.09451.10572.90463.52904.12244.26753.94714.60762.19462.15932.1824
C52.50961.54403.81033.14864.28834.28832.15812.19474.60464.16042.69353.82931.09401.09311.09603.96574.35795.35105.00264.12014.9642
C62.50963.14861.54403.81034.28834.28832.69353.82932.15812.19474.60464.16045.00264.12014.96421.09401.09311.09603.96574.35795.3510
C72.50963.81033.14861.54404.28834.28834.60464.16042.69353.82932.15812.19473.96574.35795.35105.00264.12014.96421.09401.09311.0960
H82.10701.09452.53063.38592.15812.69354.60461.77313.59382.62963.59383.74293.07892.49002.47942.11623.20043.61234.88524.68265.5013
H92.16961.10572.89532.64572.19473.82934.16041.77313.74292.65892.62962.65892.55013.08402.53913.59804.39664.65944.62094.52994.7901
H102.10703.38591.09452.53064.60462.15812.69353.59383.74291.77313.59382.62964.88524.68265.50133.07892.49002.47942.11623.20043.6123
H112.16962.64571.10572.89534.16042.19473.82932.62962.65891.77313.74292.65894.62094.52994.79012.55013.08402.53913.59804.39664.6594
H122.10702.53063.38591.09452.69354.60462.15813.59382.62963.59383.74291.77312.11623.20043.61234.88524.68265.50133.07892.49002.4794
H132.16962.89532.64571.10573.82934.16042.19473.74292.65892.62962.65891.77313.59804.39664.65944.62094.52994.79012.55013.08402.5391
H142.81792.19464.26752.90461.09405.00263.96573.07892.55014.88524.62092.11623.59801.77621.77414.85095.01966.05624.85093.82154.4330
H152.69872.15933.94713.52901.09314.12014.35792.49003.08404.68264.52993.20044.39661.77621.78703.82153.93605.21265.01963.93605.1300
H163.45832.18244.60764.12241.09604.96425.35102.47942.53915.50134.79013.61234.65941.77411.78704.43305.13005.98636.05625.21265.9863
H172.81792.90462.19464.26753.96571.09405.00262.11623.59803.07892.55014.88524.62094.85093.82154.43301.77621.77414.85095.01966.0562
H182.69873.52902.15933.94714.35791.09314.12013.20044.39662.49003.08404.68264.52995.01963.93605.13001.77621.78703.82153.93605.2126
H193.45834.12242.18244.60765.35101.09604.96423.61234.65942.47942.53915.50134.79016.05625.21265.98631.77411.78704.43305.13005.9863
H202.81794.26752.90462.19465.00263.96571.09404.88524.62092.11623.59803.07892.55014.85095.01966.05624.85093.82154.43301.77621.7741
H212.69873.94713.52902.15934.12014.35791.09314.68264.52993.20044.39662.49003.08403.82153.93605.21265.01963.93605.13001.77621.7870
H223.45834.60764.12242.18244.96425.35101.09605.50134.79013.61234.65942.47942.53914.43305.13005.98636.05625.21265.98631.77411.7870

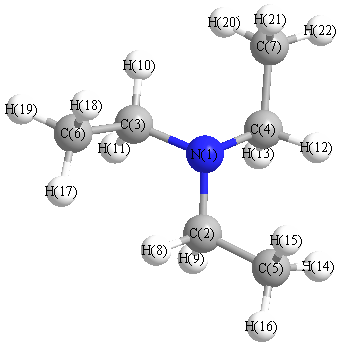
picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 110.910 N1 C2 H8 107.402
N1 C2 H9 111.658 N1 C3 C6 110.910
N1 C3 H10 107.401 N1 C3 H11 111.658
N1 C4 C7 110.910 N1 C4 H12 107.402
N1 C4 H13 111.658 C2 N1 C3 110.826
C2 N1 C4 110.826 C2 C5 H14 111.448
C2 C5 H15 108.726 C2 C5 H16 110.367
C3 N1 C4 110.826 C3 C6 H17 111.448
C3 C6 H18 108.726 C3 C6 H19 110.367
C4 C7 H20 111.448 C4 C7 H21 108.726
C4 C7 H22 110.367 C5 C2 H8 108.557
C5 C2 H9 110.750 C6 C3 H10 108.557
C6 C3 H11 110.750 C7 C4 H12 108.557
C7 C4 H13 110.750 H8 C2 H9 107.392
H10 C3 H11 107.392 H12 C4 H13 107.392
H14 C5 H15 108.611 H14 C5 H16 108.212
H15 C5 H16 109.442 H17 C6 H18 108.611
H17 C6 H19 108.212 H18 C6 H19 109.442
H20 C7 H21 108.611 H20 C7 H22 108.212
H21 C7 H22 109.442
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability