return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,4-Dioxane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-304.722106
Energy at 298.15K-304.733200
HF Energy-304.124876
Nuclear repulsion energy258.727464
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3158 3004 0.00      
2 Ag 3061 2912 0.00      
3 Ag 1583 1506 0.00      
4 Ag 1457 1386 0.00      
5 Ag 1338 1273 0.00      
6 Ag 1166 1109 0.00      
7 Ag 997 949 0.00      
8 Ag 823 783 0.00      
9 Ag 433 412 0.00      
10 Ag 394 375 0.00      
11 Au 3155 3001 54.87      
12 Au 3059 2910 29.94      
13 Au 1567 1491 0.12      
14 Au 1424 1355 27.95      
15 Au 1303 1240 12.43      
16 Au 1126 1071 7.97      
17 Au 1096 1043 92.35      
18 Au 884 841 19.76      
19 Au 236 224 2.21      
20 Bg 3157 3004 0.00      
21 Bg 3066 2916 0.00      
22 Bg 1568 1492 0.00      
23 Bg 1407 1339 0.00      
24 Bg 1256 1195 0.00      
25 Bg 1099 1046 0.00      
26 Bg 884 841 0.00      
27 Bg 463 441 0.00      
28 Bu 3160 3006 52.66      
29 Bu 3068 2918 108.52      
30 Bu 1579 1502 10.35      
31 Bu 1469 1398 0.00      
32 Bu 1328 1263 3.40      
33 Bu 1082 1030 8.96      
34 Bu 826 786 30.91      
35 Bu 591 562 17.64      
36 Bu 269 256 26.38      

Unscaled Zero Point Vibrational Energy (zpe) 27265.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 25937.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.16088 0.15100 0.08810

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.767 1.201
C2 0.000 0.767 -1.201
C3 0.000 -0.767 1.201
C4 0.000 -0.767 -1.201
O5 -0.709 -1.265 0.000
O6 0.709 1.265 0.000
H7 -1.039 1.125 1.209
H8 -1.039 1.125 -1.209
H9 1.039 -1.125 -1.209
H10 1.039 -1.125 1.209
H11 0.544 1.170 -2.059
H12 0.544 1.170 2.059
H13 -0.544 -1.170 -2.059
H14 -0.544 -1.170 2.059

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13 H14
C12.40241.53362.85012.46431.48111.09892.64893.23532.15833.33011.09283.83102.1868
C22.40242.85011.53362.46431.48112.64891.09892.15833.23531.09283.33012.18683.8310
C31.53362.85012.40241.48112.46432.15833.23532.64891.09893.83102.18683.33011.0928
C42.85011.53362.40241.48112.46433.23532.15831.09892.64892.18683.83101.09283.3301
O52.46432.46431.48111.48112.90002.69832.69832.13002.13003.42593.42592.06832.0683
O61.48111.48112.46432.46432.90002.13002.13002.69832.69832.06832.06833.42593.4259
H71.09892.64892.15833.23532.69832.13002.41803.90213.06263.63161.79704.02412.4968
H82.64891.09893.23532.15832.69832.13002.41803.06263.90211.79703.63162.49684.0241
H93.23532.15832.64891.09892.13002.69833.90213.06262.41802.49684.02411.79703.6316
H102.15833.23531.09892.64892.13002.69833.06263.90212.41804.02412.49683.63161.7970
H113.33011.09283.83102.18683.42592.06833.63161.79702.49684.02414.11892.57964.8600
H121.09283.33012.18683.83103.42592.06831.79703.63164.02412.49684.11894.86002.5796
H133.83102.18683.33011.09282.06833.42594.02412.49681.79703.63162.57964.86004.1189
H142.18683.83101.09283.33012.06833.42592.49684.02413.63161.79704.86002.57964.1189

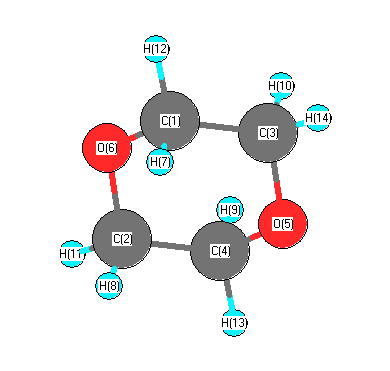
picture of 1,4-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 O5 109.644 C1 C3 H10 109.022
C1 C3 H14 111.632 C1 O6 C2 108.388
C2 C4 O5 109.644 C2 C4 H9 109.022
C2 C4 H13 111.632 C3 C1 O6 109.644
C3 C1 H7 109.022 C3 C1 H12 111.632
C3 O5 C4 108.388 C4 C2 O6 109.644
C4 C2 H8 109.022 C4 C2 H11 111.632
O5 C3 H10 110.417 O5 C3 H14 105.953
O5 C4 H9 110.417 O5 C4 H13 105.953
O6 C1 H7 110.417 O6 C1 H12 105.953
O6 C2 H8 110.417 O6 C2 H11 105.953
H7 C1 H12 110.147 H8 C2 H11 110.147
H9 C4 H13 110.147 H10 C3 H14 110.147
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability