return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (1-Butanol, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-270.260848
Energy at 298.15K-270.274403
HF Energy-269.680041
Nuclear repulsion energy256.219364
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3513 3342 0.36      
2 A 3195 3039 8.78      
3 A 3162 3008 40.79      
4 A 3151 2998 32.51      
5 A 3144 2991 33.56      
6 A 3112 2960 19.00      
7 A 3095 2944 47.66      
8 A 3084 2934 25.59      
9 A 3077 2927 2.57      
10 A 3075 2925 32.35      
11 A 3067 2917 12.37      
12 A 3028 2880 35.84      
13 A 1624 1545 2.35      
14 A 1613 1535 7.01      
15 A 1608 1530 6.20      
16 A 1603 1525 5.46      
17 A 1594 1517 7.91      
18 A 1585 1508 1.85      
19 A 1498 1425 2.60      
20 A 1491 1419 9.28      
21 A 1470 1399 5.78      
22 A 1446 1376 1.01      
23 A 1413 1344 0.71      
24 A 1397 1329 0.69      
25 A 1363 1296 5.70      
26 A 1309 1246 1.79      
27 A 1288 1225 28.91      
28 A 1228 1168 0.91      
29 A 1204 1145 7.52      
30 A 1156 1099 10.44      
31 A 1066 1014 0.93      
32 A 1048 997 5.44      
33 A 1023 973 23.85      
34 A 1016 966 45.20      
35 A 973 926 1.76      
36 A 920 875 1.45      
37 A 846 805 3.68      
38 A 785 747 8.39      
39 A 505 480 5.88      
40 A 449 427 2.29      
41 A 405 385 2.72      
42 A 298 284 7.58      
43 A 275 262 74.69      
44 A 265 253 46.43      
45 A 236 224 11.12      
46 A 231 220 1.09      
47 A 128 121 3.95      
48 A 91 87 4.24      

Unscaled Zero Point Vibrational Energy (zpe) 37074.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35268.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.16733 0.07359 0.05573

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
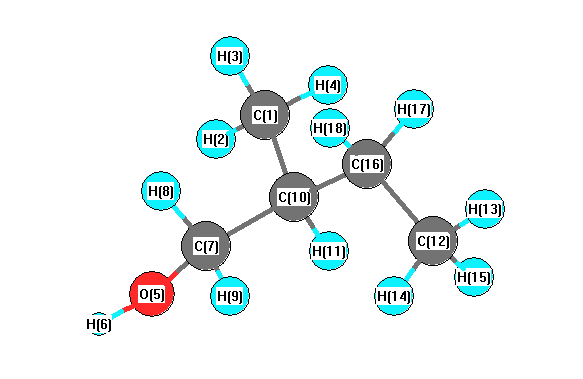
C1 -0.824 1.661 -0.006
H2 -1.857 1.526 -0.331
H3 -0.814 1.881 1.070
H4 -0.374 2.506 -0.539
O5 -2.120 -0.842 -0.240
H6 -2.641 -1.560 0.206
C7 -0.773 -0.827 0.363
H8 -0.815 -0.670 1.452
H9 -0.241 -1.768 0.168
C10 -0.034 0.358 -0.278
H11 -0.010 0.178 -1.362
C12 2.338 -0.710 -0.176
H13 3.365 -0.538 0.165
H14 1.995 -1.658 0.249
H15 2.347 -0.804 -1.268
C16 1.420 0.467 0.247
H17 1.850 1.404 -0.131
H18 1.393 0.546 1.343

Atom - Atom Distances (Å)
  C1 H2 H3 H4 O5 H6 C7 H8 H9 C10 H11 C12 H13 H14 H15 C16 H17 H18
C11.09171.09781.09582.82853.70362.51542.74903.48191.54782.16773.95594.73434.36174.21062.55432.68922.8246
H21.09171.78221.79002.38443.22822.68173.01443.70232.16602.50834.75665.63745.03104.89793.49233.71473.7853
H31.09781.78221.78113.29193.99002.79882.57903.80172.17843.07524.26644.91304.59224.76162.76882.96102.5940
H41.09581.79001.78113.78784.71393.47553.77394.33342.18982.49524.22234.87244.85474.34682.82692.51493.2414
O52.82852.38443.29193.78780.99231.47562.14432.13382.40742.59884.46065.50874.22314.58453.80604.56294.0958
H63.70363.22823.99004.71390.99232.01242.38342.40923.27273.52165.06556.09274.63685.25624.53915.39134.6904
C72.51542.68172.79883.47551.47562.01241.10161.09831.53722.13723.16004.15342.89233.52142.54953.47902.7457
H82.74903.01442.57903.77392.14432.38341.10161.78462.15933.04733.54904.37603.21224.17372.78213.72972.5226
H93.48193.70233.80174.33342.13382.40921.09831.78462.18212.48582.80883.81032.23983.11292.78593.81083.0666
C101.54782.16602.17842.18982.40743.27271.53722.15932.18211.09812.60333.54302.90832.82871.54982.15962.1682
H112.16772.50833.07522.49522.59883.52162.13723.04732.48581.09812.77603.77253.15962.55552.17162.54433.0690
C123.95594.75664.26644.22234.46065.06553.16003.54902.80882.60332.77601.09581.09451.09591.55192.17032.1863
H134.73435.63744.91304.87245.50876.09274.15344.37603.81033.54303.77251.09581.77231.77742.19122.48102.5405
H144.36175.03104.59224.85474.22314.63682.89233.21222.23982.90833.15961.09451.77231.77612.20193.08932.5333
H154.21064.89794.76164.34684.58455.25623.52144.17373.11292.82872.55551.09591.77741.77612.18492.53333.0910
C162.55433.49232.76882.82693.80604.53912.54952.78212.78591.54982.17161.55192.19122.20192.18491.09811.0993
H172.68923.71472.96102.51494.56295.39133.47903.72973.81082.15962.54432.17032.48103.08932.53331.09811.7666
H182.82463.78532.59403.24144.09584.69042.74572.52263.06662.16823.06902.18632.54052.53333.09101.09931.7666

picture of 1-Butanol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C10 C7 109.246 C1 C10 H11 108.837
C1 C10 C16 111.094 H2 C1 H3 108.973
H2 C1 H4 109.827 H2 C1 C10 109.069
H3 C1 H4 108.570 H3 C1 C10 109.682
H4 C1 C10 110.695 O5 C7 H8 111.792
O5 C7 H9 111.144 O5 C7 C10 106.062
H6 O5 C7 107.640 C7 C10 H11 107.206
C7 C10 C16 111.352 H8 C7 H9 108.435
H8 C7 C10 108.707 H9 C7 C10 110.676
C10 C16 C12 114.134 C10 C16 H17 108.076
C10 C16 H18 108.671 H11 C10 C16 108.997
C12 C16 H17 108.758 C12 C16 H18 109.929
H13 C12 H14 108.032 H13 C12 H15 108.380
H13 C12 C16 110.515 H14 C12 H15 108.359
H14 C12 C16 111.445 H15 C12 C16 110.018
H17 C16 H18 107.023
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability