Jump to
S1C2
Energy calculated at M06-2X/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1190.089568 |
| Energy at 298.15K | -1190.092041 |
| HF Energy | -1190.089568 |
| Nuclear repulsion energy | 196.155740 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2711 |
2568 |
0.57 |
|
|
|
| 2 |
A |
955 |
904 |
1.55 |
|
|
|
| 3 |
A |
489 |
463 |
0.38 |
|
|
|
| 4 |
A |
343 |
325 |
31.03 |
|
|
|
| 5 |
A |
209 |
198 |
0.00 |
|
|
|
| 6 |
B |
2711 |
2568 |
5.17 |
|
|
|
| 7 |
B |
934 |
885 |
10.42 |
|
|
|
| 8 |
B |
471 |
447 |
27.63 |
|
|
|
| 9 |
B |
406 |
385 |
21.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4614.5 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 4370.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.849 |
| S2 |
0.000 |
1.652 |
-0.387 |
| S3 |
0.000 |
-1.652 |
-0.387 |
| H4 |
-1.324 |
1.732 |
-0.607 |
| H5 |
1.324 |
-1.732 |
-0.607 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0631 | 2.0631 | 2.6218 | 2.6218 |
S2 | 2.0631 | | 3.3040 | 1.3446 | 3.6404 | S3 | 2.0631 | 3.3040 | | 3.6404 | 1.3446 | H4 | 2.6218 | 1.3446 | 3.6404 | | 4.3601 | H5 | 2.6218 | 3.6404 | 1.3446 | 4.3601 | |
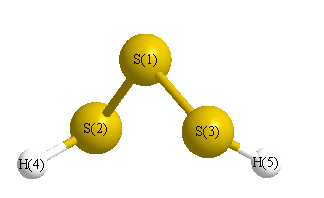 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.393 |
|
S1 |
S3 |
H5 |
98.393 |
| S2 |
S1 |
S3 |
106.401 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.048 |
|
|
|
| 2 |
S |
-0.135 |
|
|
|
| 3 |
S |
-0.135 |
|
|
|
| 4 |
H |
0.159 |
|
|
|
| 5 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.717 |
0.717 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.668 |
-4.372 |
0.000 |
| y |
-4.372 |
-40.351 |
0.000 |
| z |
0.000 |
0.000 |
-41.982 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.498 |
-4.372 |
0.000 |
| y |
-4.372 |
-0.526 |
0.000 |
| z |
0.000 |
0.000 |
-2.972 |
|
| Polar |
|---|
| 3z2-r2 | -5.945 |
|---|
| x2-y2 | 2.683 |
|---|
| xy | -4.372 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.116 |
-0.683 |
0.000 |
| y |
-0.683 |
10.195 |
0.000 |
| z |
0.000 |
0.000 |
5.457 |
<r2> (average value of r
2) Å
2
| <r2> |
138.877 |
| (<r2>)1/2 |
11.785 |
Jump to
S1C1
Energy calculated at M06-2X/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1190.089008 |
| Energy at 298.15K | |
| HF Energy | -1190.089008 |
| Nuclear repulsion energy | 196.182055 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2568 |
2432 |
6.77 |
|
|
|
| 2 |
A' |
948 |
898 |
8.36 |
|
|
|
| 3 |
A' |
488 |
462 |
0.64 |
|
|
|
| 4 |
A' |
213 |
201 |
1.73 |
|
|
|
| 5 |
A' |
86 |
82 |
21.75 |
|
|
|
| 6 |
A" |
2569 |
2434 |
0.30 |
|
|
|
| 7 |
A" |
940 |
890 |
7.48 |
|
|
|
| 8 |
A" |
472 |
447 |
25.00 |
|
|
|
| 9 |
A" |
150i |
142i |
14.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4067.1 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 3852.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.053 |
0.845 |
0.000 |
| S2 |
-0.053 |
-0.390 |
1.652 |
| S3 |
-0.053 |
-0.390 |
-1.652 |
| H4 |
1.276 |
-0.525 |
1.797 |
| H5 |
1.276 |
-0.525 |
-1.797 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0620 | 2.0620 | 2.6214 | 2.6214 |
S2 | 2.0620 | | 3.3036 | 1.3443 | 3.6985 | S3 | 2.0620 | 3.3036 | | 3.6985 | 1.3443 | H4 | 2.6214 | 1.3443 | 3.6985 | | 3.5935 | H5 | 2.6214 | 3.6985 | 1.3443 | 3.5935 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.425 |
|
S1 |
S3 |
H5 |
98.425 |
| S2 |
S1 |
S3 |
106.460 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.048 |
|
|
|
| 2 |
S |
-0.126 |
|
|
|
| 3 |
S |
-0.126 |
|
|
|
| 4 |
H |
0.151 |
|
|
|
| 5 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.161 |
-0.569 |
0.000 |
2.235 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.912 |
-1.624 |
0.000 |
| y |
-1.624 |
-42.207 |
0.000 |
| z |
0.000 |
0.000 |
-39.962 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.172 |
-1.624 |
0.000 |
| y |
-1.624 |
-3.270 |
0.000 |
| z |
0.000 |
0.000 |
0.098 |
|
| Polar |
|---|
| 3z2-r2 | 0.196 |
|---|
| x2-y2 | 4.295 |
|---|
| xy | -1.624 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.090 |
-0.372 |
0.000 |
| y |
-0.372 |
5.392 |
0.000 |
| z |
0.000 |
0.000 |
10.260 |
<r2> (average value of r
2) Å
2
| <r2> |
138.986 |
| (<r2>)1/2 |
11.789 |
