return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH2OH (hydroxy acetaldehyde)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS H in 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-226.907180
Energy at 298.15K-226.912153
HF Energy-226.479036
Nuclear repulsion energy117.299051
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3433 3266 24.39      
2 A' 3062 2913 48.19      
3 A' 3041 2893 48.20      
4 A' 1632 1553 9.27      
5 A' 1593 1516 13.64      
6 A' 1489 1416 110.76      
7 A' 1439 1369 8.50      
8 A' 1330 1265 48.08      
9 A' 1049 998 32.69      
10 A' 839 798 42.86      
11 A' 758 721 7.44      
12 A' 319 304 21.92      
13 A" 3086 2936 29.10      
14 A" 1259 1198 7.99      
15 A" 1150 1094 0.17      
16 A" 803 764 0.09      
17 A" 455 433 105.52      
18 A" 187 178 9.97      

Unscaled Zero Point Vibrational Energy (zpe) 13462.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12806.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.56723 0.21829 0.16246

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.973 0.000
C2 0.955 -0.221 0.000
O3 -1.378 0.497 0.000
O4 0.517 -1.396 0.000
H5 -1.294 -0.499 0.000
H6 0.209 1.586 0.888
H7 0.209 1.586 -0.888
H8 2.029 0.008 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 O4 H5 H6 H7 H8
C11.52871.45772.42481.95911.09891.09892.2467
C21.52872.44091.25442.26592.14682.14681.0980
O31.45772.44092.67870.99912.11922.11923.4418
O42.42481.25442.67872.02113.12653.12652.0634
H51.95912.26590.99912.02112.71832.71833.3610
H61.09892.14682.11923.12652.71831.77592.5672
H71.09892.14682.11923.12652.71831.77592.5672
H82.24671.09803.44182.06343.36102.56722.5672

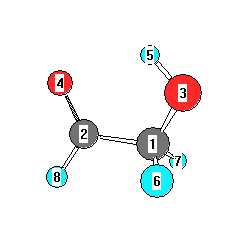
picture of hydroxy acetaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 120.896 C1 C2 H8 116.638
C1 O3 H5 104.215 C2 C1 O3 109.617
C2 C1 H6 108.467 C2 C1 H7 108.467
O3 C1 H6 111.197 O3 C1 H7 111.197
O4 C2 H8 122.466 H6 C1 H7 107.807
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability