return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,3-Hexadiene, (Z)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.228925
Energy at 298.15K-232.238779
HF Energy-231.687930
Nuclear repulsion energy215.088014
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3267 3108 19.77      
2 A' 3243 3085 0.42      
3 A' 3167 3012 5.74      
4 A' 3152 2999 33.63      
5 A' 3148 2995 33.74      
6 A' 3119 2968 3.65      
7 A' 3086 2936 14.83      
8 A' 3057 2908 17.92      
9 A' 1695 1613 1.97      
10 A' 1646 1566 0.79      
11 A' 1612 1533 7.12      
12 A' 1578 1501 2.59      
13 A' 1535 1460 10.92      
14 A' 1492 1420 4.35      
15 A' 1468 1397 0.66      
16 A' 1432 1362 2.97      
17 A' 1386 1318 0.15      
18 A' 1369 1302 4.74      
19 A' 1203 1145 0.13      
20 A' 1115 1061 4.10      
21 A' 1060 1009 3.01      
22 A' 936 891 1.49      
23 A' 795 756 1.92      
24 A' 729 694 1.68      
25 A' 436 415 0.16      
26 A' 286 272 1.66      
27 A' 205 195 0.32      
28 A" 3176 3022 22.56      
29 A" 3084 2934 14.93      
30 A" 1607 1529 5.51      
31 A" 1375 1308 0.06      
32 A" 1172 1115 2.99      
33 A" 1039 989 16.11      
34 A" 999 951 6.92      
35 A" 942 896 43.88      
36 A" 862 820 1.30      
37 A" 781 743 7.67      
38 A" 627 597 32.60      
39 A" 395 376 0.14      
40 A" 313 297 0.03      
41 A" 86 82 0.17      
42 A" 44 41 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 31860.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30308.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.22615 0.06688 0.05264

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.067 1.288 0.000
C2 -1.238 0.576 0.000
H3 0.045 2.215 0.000
C4 0.000 1.122 0.000
H5 1.390 -0.601 0.000
C6 1.329 0.481 0.000
H7 3.448 0.724 0.000
H8 2.462 2.287 0.000
C9 2.472 1.199 0.000
H10 -2.385 -0.990 0.879
H11 -2.385 -0.990 -0.879
C12 -1.734 -0.868 0.000
H13 -1.212 -2.978 0.000
H14 -0.059 -1.969 -0.891
H15 -0.059 -1.969 0.891
C16 -0.691 -2.013 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 C6 H7 H8 C9 H10 H11 C12 H13 H14 H15 C16
H11.09222.30652.07373.93923.49085.54424.63814.54022.46192.46192.18074.35023.92863.92863.5761
C21.09222.08131.35352.87982.56934.68924.07693.76262.13082.13081.52643.55402.94342.94342.6465
H32.30652.08131.09333.12052.15793.71562.41822.63124.11684.11683.55895.34264.27904.27904.2915
C42.07371.35351.09332.21401.47603.47132.72372.47343.30513.30512.63964.27573.21803.21803.2108
H53.93922.87983.12052.21401.08332.44823.08092.10053.89553.89553.13563.52442.18302.18302.5150
C63.49082.56932.15791.47601.08332.13322.13251.35014.09054.09053.34714.29192.95342.95343.2095
H75.54424.68923.71563.47132.44822.13321.84831.08566.14356.14355.42175.95224.51124.51124.9629
H84.63814.07692.41822.72373.08092.13251.84831.08825.91685.91685.25006.42055.02665.02665.3326
C94.54023.76262.63122.47342.10051.35011.08561.08825.40005.40004.68695.56994.15214.15214.5085
H102.46192.13084.11683.30513.89554.09056.14355.91685.40001.75701.10022.46953.08272.52402.1653
H112.46192.13084.11683.30513.89554.09056.14355.91685.40001.75701.10022.46952.52403.08272.1653
C122.18071.52643.55892.63963.13563.34715.42175.25004.68691.10021.10022.17392.19442.19441.5495
H134.35023.55405.34264.27573.52444.29195.95226.42055.56992.46952.46952.17391.77271.77271.0964
H143.92862.94344.27903.21802.18302.95344.51125.02664.15213.08272.52402.19441.77271.78271.0937
H153.92862.94344.27903.21802.18302.95344.51125.02664.15212.52403.08272.19441.77271.78271.0937
C163.57612.64654.29153.21082.51503.20954.96295.33264.50852.16532.16531.54951.09641.09371.0937

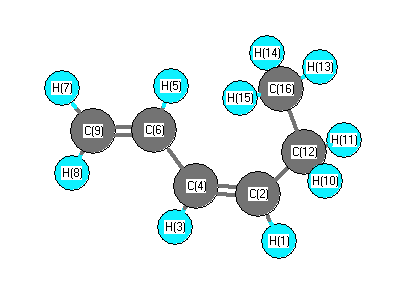
picture of 1,3-Hexadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 115.550 H1 C2 C12 111.687
C2 C4 H3 116.151 C2 C4 C6 130.427
C2 C12 H10 107.312 C2 C12 H11 107.312
C2 C12 C16 118.719 H3 C4 C6 113.422
C4 C2 C12 132.762 C4 C6 H5 118.985
C4 C6 C9 122.073 H5 C6 C9 118.942
C6 C9 H7 121.902 C6 C9 H8 121.616
H7 C9 H8 116.482 H10 C12 H11 105.971
H10 C12 C16 108.418 H11 C12 C16 108.418
C12 C16 H13 109.296 C12 C16 H14 111.063
C12 C16 H15 111.063 H13 C16 H14 108.074
H13 C16 H15 108.074 H14 C16 H15 109.175
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability