Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -688.173069 |
| Energy at 298.15K | -688.172338 |
| HF Energy | -687.952155 |
| Nuclear repulsion energy | 59.542908 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.338 |
| C2 |
0.000 |
0.000 |
-1.307 |
| N3 |
0.000 |
0.000 |
-2.512 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.6456 | 3.8497 |
C2 | 2.6456 | | 1.2041 | N3 | 3.8497 | 1.2041 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -688.175907 |
| Energy at 298.15K | |
| HF Energy | -687.965585 |
| Nuclear repulsion energy | 63.270061 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.230 |
| C2 |
0.000 |
0.000 |
-2.449 |
| N3 |
0.000 |
0.000 |
-1.240 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6794 | 2.4706 |
C2 | 3.6794 | | 1.2088 | N3 | 2.4706 | 1.2088 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -688.183695 |
| Energy at 298.15K | -688.183581 |
| HF Energy | -687.963884 |
| Nuclear repulsion energy | 67.174754 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.060 |
0.000 |
| C2 |
0.648 |
-1.640 |
0.000 |
| N3 |
-0.555 |
-1.472 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7763 | 2.5924 |
C2 | 2.7763 | | 1.2147 | N3 | 2.5924 | 1.2147 | |
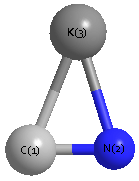 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
68.585 |
|
K1 |
N3 |
C2 |
85.552 |
| C2 |
K1 |
N3 |
25.863 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
