Jump to
S1C2
S1C3
Energy calculated at B97D3/3-21G*
| | hartrees |
|---|
| Energy at 0K | -689.255217 |
| Energy at 298.15K | -689.254362 |
| HF Energy | -689.255217 |
| Nuclear repulsion energy | 60.299948 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B97D3/3-21G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.322 |
| C2 |
0.000 |
0.000 |
-1.291 |
| N3 |
0.000 |
0.000 |
-2.480 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.6129 | 3.8015 |
C2 | 2.6129 | | 1.1886 | N3 | 3.8015 | 1.1886 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.538 |
|
|
|
| 2 |
C |
-0.085 |
|
|
|
| 3 |
N |
-0.453 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
11.804 |
11.804 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.025 |
0.000 |
0.000 |
| y |
0.000 |
-24.025 |
0.000 |
| z |
0.000 |
0.000 |
-29.102 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.539 |
0.000 |
0.000 |
| y |
0.000 |
2.539 |
0.000 |
| z |
0.000 |
0.000 |
-5.077 |
|
| Polar |
|---|
| 3z2-r2 | -10.155 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.144 |
0.000 |
0.000 |
| y |
0.000 |
4.144 |
0.000 |
| z |
0.000 |
0.000 |
8.018 |
<r2> (average value of r
2) Å
2
| <r2> |
102.304 |
| (<r2>)1/2 |
10.115 |
Jump to
S1C1
S1C3
Energy calculated at B97D3/3-21G*
| | hartrees |
|---|
| Energy at 0K | -689.261577 |
| Energy at 298.15K | |
| HF Energy | -689.261577 |
| Nuclear repulsion energy | 63.810066 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B97D3/3-21G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.220 |
| C2 |
0.000 |
0.000 |
-2.428 |
| N3 |
0.000 |
0.000 |
-1.230 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6482 | 2.4498 |
C2 | 3.6482 | | 1.1985 | N3 | 2.4498 | 1.1985 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.516 |
|
|
|
| 2 |
C |
0.007 |
|
|
|
| 3 |
N |
-0.524 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
11.838 |
11.838 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.036 |
0.000 |
0.000 |
| y |
0.000 |
-24.036 |
0.000 |
| z |
0.000 |
0.000 |
-31.555 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.760 |
0.000 |
0.000 |
| y |
0.000 |
3.760 |
0.000 |
| z |
0.000 |
0.000 |
-7.519 |
|
| Polar |
|---|
| 3z2-r2 | -15.038 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.036 |
0.000 |
0.000 |
| y |
0.000 |
4.036 |
0.000 |
| z |
0.000 |
0.000 |
8.067 |
<r2> (average value of r
2) Å
2
| <r2> |
90.822 |
| (<r2>)1/2 |
9.530 |
Jump to
S1C1
S1C2
Energy calculated at B97D3/3-21G*
| | hartrees |
|---|
| Energy at 0K | -689.265529 |
| Energy at 298.15K | -689.265330 |
| HF Energy | -689.265529 |
| Nuclear repulsion energy | 67.293245 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B97D3/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.066 |
0.000 |
| C2 |
0.629 |
-1.709 |
0.000 |
| N3 |
-0.539 |
-1.429 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.8448 | 2.5522 |
C2 | 2.8448 | | 1.2007 | N3 | 2.5522 | 1.2007 | |
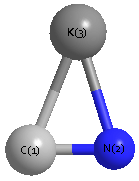 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
63.758 |
|
K1 |
N3 |
C2 |
91.284 |
| C2 |
K1 |
N3 |
24.958 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.467 |
|
|
|
| 2 |
C |
-0.008 |
|
|
|
| 3 |
N |
-0.460 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.369 |
10.252 |
0.000 |
10.259 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.296 |
1.237 |
0.000 |
| y |
1.237 |
-25.867 |
0.000 |
| z |
0.000 |
0.000 |
-24.186 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.270 |
1.237 |
0.000 |
| y |
1.237 |
0.375 |
0.000 |
| z |
0.000 |
0.000 |
2.895 |
|
| Polar |
|---|
| 3z2-r2 | 5.790 |
|---|
| x2-y2 | -2.429 |
|---|
| xy | 1.237 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.886 |
-0.218 |
0.000 |
| y |
-0.218 |
4.749 |
0.000 |
| z |
0.000 |
0.000 |
3.831 |
<r2> (average value of r
2) Å
2
| <r2> |
74.107 |
| (<r2>)1/2 |
8.609 |
