return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (Cyclopropane, 1,1-dimethyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-194.502089
Energy at 298.15K-194.512961
HF Energy-194.043957
Nuclear repulsion energy183.060982
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3190 3034 6.64      
2 A 3144 2991 33.15      
3 A 3068 2919 22.29      
4 A 1607 1529 3.42      
5 A 1572 1495 2.25      
6 A 1499 1426 0.04      
7 A 1365 1299 0.82      
8 A 1126 1072 20.33      
9 A 1093 1039 0.33      
10 A 926 881 10.14      
11 A 664 632 1.75      
12 A 363 345 0.00      
13 A 3270 3111 0.00      
14 A 3135 2982 0.00      
15 A 1585 1508 0.00      
16 A 1226 1167 0.00      
17 A 1123 1068 0.00      
18 A 947 901 0.00      
19 A 342 326 0.00      
20 A 230 219 0.00      
21 A 3184 3029 12.50      
22 A 3138 2985 48.47      
23 A 1601 1523 5.94      
24 A 1554 1479 3.72      
25 A 1232 1172 7.13      
26 A 1137 1082 1.24      
27 A 837 796 5.57      
28 A 398 378 1.96      
29 A 246 234 0.03      
30 A 3285 3125 15.93      
31 A 3141 2988 12.56      
32 A 3066 2916 25.50      
33 A 1597 1519 5.13      
34 A 1484 1412 8.42      
35 A 1351 1286 1.94      
36 A 1032 982 0.26      
37 A 968 921 1.05      
38 A 821 781 0.02      
39 A 363 345 0.61      

Unscaled Zero Point Vibrational Energy (zpe) 30954.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 29447.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.20194 0.17024 0.12528

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.156
C2 0.770 0.000 -1.164
C3 -0.770 0.000 -1.164
C4 0.000 1.284 0.978
C5 0.000 -1.284 0.978
H6 1.275 0.915 -1.455
H7 1.275 -0.915 -1.455
H8 -1.275 -0.915 -1.455
H9 -1.275 0.915 -1.455
H10 -0.891 1.328 1.617
H11 -0.891 -1.328 1.617
H12 0.891 -1.328 1.617
H13 0.891 1.328 1.617
H14 0.000 2.164 0.323
H15 0.000 -2.164 0.323

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52871.52871.52481.52482.24912.24912.24912.24912.16552.16552.16552.16552.16992.1699
C21.52871.53952.61392.61391.08461.08462.25862.25863.50083.50083.08433.08432.73592.7359
C31.52871.53952.61392.61392.25862.25861.08461.08463.08433.08433.50083.50082.73592.7359
C41.52482.61392.61392.56892.77163.51883.51882.77161.09702.83272.83271.09701.09643.5097
C51.52482.61392.61392.56893.51882.77162.77163.51882.83271.09701.09702.83273.50971.0964
H62.24911.08462.25862.77163.51881.82963.13792.54933.78114.37663.82273.12332.51893.7765
H72.24911.08462.25863.51882.77161.82962.54933.13794.37663.78113.12333.82273.77652.5189
H82.24912.25861.08463.51882.77163.13792.54931.82963.82273.12333.78114.37663.77652.5189
H92.24912.25861.08462.77163.51882.54933.13791.82963.12333.82274.37663.78112.51893.7765
H102.16553.50083.08431.09702.83273.78114.37663.82273.12332.65533.19751.78141.77963.8284
H112.16553.50083.08432.83271.09704.37663.78113.12333.82272.65531.78143.19753.82841.7796
H122.16553.08433.50082.83271.09703.82273.12333.78114.37663.19751.78142.65533.82841.7796
H132.16553.08433.50081.09702.83273.12333.82274.37663.78111.78143.19752.65531.77963.8284
H142.16992.73592.73591.09643.50972.51893.77653.77652.51891.77963.82843.82841.77964.3270
H152.16992.73592.73593.50971.09643.77652.51892.51893.77653.82841.77961.77963.82844.3270

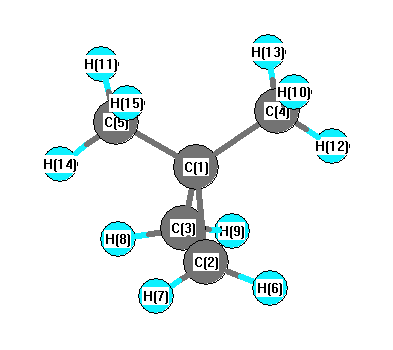
picture of Cyclopropane, 1,1-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 59.764 C1 C2 H6 117.776
C1 C2 H7 117.776 C1 C3 C2 59.764
C1 C3 H8 117.776 C1 C3 H9 117.776
C1 C4 H10 110.305 C1 C4 H13 110.305
C1 C4 H14 110.687 C1 C5 H11 110.305
C1 C5 H12 110.305 C1 C5 H15 110.687
C2 C1 C3 60.472 C2 C1 C4 117.750
C2 C1 C5 117.750 C2 C3 H8 117.744
C2 C3 H9 117.744 C3 C1 C4 117.750
C3 C1 C5 117.750 C3 C2 H6 117.744
C3 C2 H7 117.744 C4 C1 C5 114.779
H6 C2 H7 115.016 H8 C3 H9 115.016
H10 C4 H13 108.578 H10 C4 H14 108.451
H11 C5 H12 108.578 H11 C5 H15 108.451
H12 C5 H15 108.451 H13 C4 H14 108.451
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability