return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (Propane, 2-methoxy-2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-270.264752
Energy at 298.15K-270.278133
HF Energy-269.680126
Nuclear repulsion energy267.564500
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3174 3020 41.14      
2 A 3167 3013 20.22      
3 A 3165 3011 10.72      
4 A 3159 3006 44.77      
5 A 3091 2941 5.27      
6 A 3085 2935 16.25      
7 A 3061 2912 27.96      
8 A 1621 1542 16.49      
9 A 1616 1537 6.76      
10 A 1598 1521 3.06      
11 A 1588 1511 1.29      
12 A 1535 1460 3.57      
13 A 1496 1423 4.80      
14 A 1470 1399 13.69      
15 A 1320 1256 18.72      
16 A 1291 1228 63.99      
17 A 1211 1152 7.75      
18 A 1105 1051 10.98      
19 A 1061 1009 57.01      
20 A 950 904 0.15      
21 A 845 804 6.74      
22 A 705 670 3.30      
23 A 502 477 2.24      
24 A 401 382 0.90      
25 A 371 353 1.04      
26 A 287 273 0.93      
27 A 257 245 0.73      
28 A 3176 3022 23.41      
29 A 3166 3012 4.51      
30 A 3156 3002 0.18      
31 A 3115 2963 59.79      
32 A 3082 2932 6.02      
33 A 1606 1528 0.15      
34 A 1592 1515 5.78      
35 A 1585 1508 0.00      
36 A 1575 1499 0.01      
37 A 1470 1398 15.41      
38 A 1305 1242 24.03      
39 A 1184 1127 0.41      
40 A 1094 1041 2.31      
41 A 1018 969 0.00      
42 A 940 894 0.54      
43 A 443 422 5.45      
44 A 346 329 2.93      
45 A 286 273 1.57      
46 A 242 230 0.74      
47 A 196 186 2.53      
48 A 60 57 4.01      

Unscaled Zero Point Vibrational Energy (zpe) 36884.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35088.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14293 0.08876 0.08817

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.799 -0.835 0.000
C2 0.000 0.418 0.000
C3 -1.104 1.492 0.000
C4 -0.018 -2.079 0.000
H5 -0.763 -2.878 0.000
H6 -0.666 2.496 0.000
C7 0.863 0.529 1.274
C8 0.863 0.529 -1.274
H9 0.614 -2.182 0.891
H10 0.614 -2.182 -0.891
H11 0.231 0.356 -2.151
H12 0.231 0.356 2.151
H13 1.302 1.530 1.337
H14 1.302 1.530 -1.337
H15 1.677 -0.204 1.271
H16 1.677 -0.204 -1.271
H17 -1.728 1.363 -0.889
H18 -1.728 1.363 0.889

Atom - Atom Distances (Å)
  O1 C2 C3 C4 H5 H6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
O11.48602.34701.46952.04413.33312.49872.49872.14562.14562.66542.66543.43383.43382.85402.85402.54632.5463
C21.48601.54012.49783.38392.18141.54281.54282.81632.81632.16422.16422.17182.17182.19482.19482.16092.1609
C32.34701.54013.73334.38421.09492.53392.53394.15264.15262.77462.77462.75202.75203.49703.49701.09401.0940
C41.46952.49783.73331.09254.62083.03323.03321.09671.09673.25883.25884.06894.06892.82962.82963.94543.9454
H52.04413.38394.38421.09255.37513.98423.98421.78131.78134.00944.00945.04825.04823.83703.83704.43994.4399
H63.33312.18141.09494.62085.37512.79822.79824.93064.93063.16353.16352.56682.56683.79403.79401.78971.7897
C72.49871.54282.53393.03323.98422.79822.54812.74883.47793.48721.09491.09532.83041.09522.77113.47712.7496
C82.49871.54282.53393.03323.98422.79822.54813.47792.74881.09493.48722.83041.09532.77111.09522.74963.4771
H92.14562.81634.15261.09671.78134.93062.74883.47791.78173.98012.85943.80164.38352.27793.11764.60644.2487
H102.14562.81634.15261.09671.78134.93063.47792.74881.78172.85943.98014.38353.80163.11762.27794.24874.6064
H112.66542.16422.77463.25884.00943.16353.48721.09493.98012.85944.30193.83261.78583.75761.78332.53853.7539
H122.66542.16422.77463.25884.00943.16351.09493.48722.85943.98014.30191.78583.83261.78333.75763.75392.5385
H133.43382.17182.75204.06895.04822.56681.09532.83043.80164.38353.83261.78582.67311.77563.15443.76313.0674
H143.43382.17182.75204.06895.04822.56682.83041.09534.38353.80161.78583.83262.67313.15441.77563.06743.7631
H152.85402.19483.49702.82963.83703.79401.09522.77112.27793.11763.75761.78331.77563.15442.54294.32673.7683
H162.85402.19483.49702.82963.83703.79402.77111.09523.11762.27791.78333.75763.15441.77562.54293.76834.3267
H172.54632.16091.09403.94544.43991.78973.47712.74964.60644.24872.53853.75393.76313.06744.32673.76831.7777
H182.54632.16091.09403.94544.43991.78972.74963.47714.24874.60643.75392.53853.06743.76313.76834.32671.7777

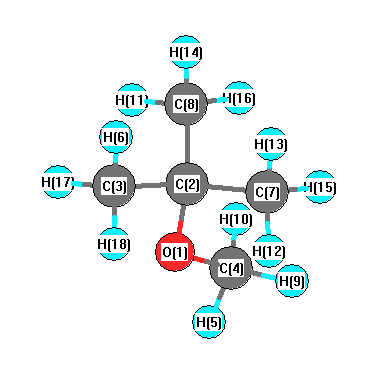
picture of Propane, 2-methoxy-2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 101.700 O1 C2 C7 111.159
O1 C2 C8 111.159 O1 C4 H5 104.895
O1 C4 H9 112.656 O1 C4 H10 112.656
C2 O1 C4 115.371 C2 C3 H6 110.622
C2 C3 H17 109.065 C2 C3 H18 109.065
C2 C7 H12 109.085 C2 C7 H13 109.660
C2 C7 H15 111.478 C2 C8 H11 109.085
C2 C8 H14 109.660 C2 C8 H16 111.478
C3 C2 C7 110.556 C3 C2 C8 110.556
H5 C4 H9 108.912 H5 C4 H10 108.912
H6 C3 H17 109.691 H6 C3 H18 109.691
C7 C2 C8 111.339 H9 C4 H10 108.653
H11 C8 H14 109.247 H11 C8 H16 109.027
H12 C7 H13 109.247 H12 C7 H15 109.027
H13 C7 H15 108.311 H14 C8 H16 108.311
H17 C3 H18 108.673
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability