Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -2599.234469 |
| Energy at 298.15K | -2599.238373 |
| HF Energy | -2599.032781 |
| Nuclear repulsion energy | 79.784632 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3221 |
3064 |
6.06 |
|
|
|
| 2 |
A' |
1483 |
1411 |
28.22 |
|
|
|
| 3 |
A' |
719 |
684 |
22.57 |
|
|
|
| 4 |
A' |
338 |
321 |
72.69 |
|
|
|
| 5 |
A" |
3374 |
3209 |
0.11 |
|
|
|
| 6 |
A" |
988 |
940 |
1.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5061.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 4815.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.007 |
1.506 |
0.000 |
| Br2 |
-0.007 |
-0.373 |
0.000 |
| H3 |
0.139 |
2.001 |
0.948 |
| H4 |
0.139 |
2.001 |
-0.948 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8787 | 1.0792 | 1.0792 |
Br2 | 1.8787 | | 2.5597 | 2.5597 | H3 | 1.0792 | 2.5597 | | 1.8961 | H4 | 1.0792 | 2.5597 | 1.8961 | |
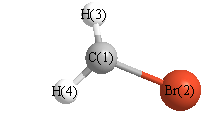 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.266 |
|
Br2 |
C1 |
H4 |
117.266 |
| H3 |
C1 |
H4 |
122.926 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -2599.234399 |
| Energy at 298.15K | |
| HF Energy | -2599.032803 |
| Nuclear repulsion energy | 79.866230 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3231 |
3074 |
4.74 |
92.25 |
0.12 |
0.21 |
| 2 |
A1 |
1481 |
1409 |
30.21 |
3.52 |
0.62 |
0.76 |
| 3 |
A1 |
722 |
687 |
20.79 |
6.35 |
0.26 |
0.41 |
| 4 |
B1 |
247i |
235i |
88.89 |
1.04 |
0.75 |
0.86 |
| 5 |
B2 |
3388 |
3223 |
0.00 |
49.71 |
0.75 |
0.86 |
| 6 |
B2 |
981 |
934 |
2.07 |
4.84 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4777.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 4545.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.503 |
| Br2 |
0.000 |
0.000 |
0.372 |
| H3 |
0.000 |
0.952 |
-2.009 |
| H4 |
0.000 |
-0.952 |
-2.009 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8754 | 1.0781 | 1.0781 |
Br2 | 1.8754 | | 2.5646 | 2.5646 | H3 | 1.0781 | 2.5646 | | 1.9041 | H4 | 1.0781 | 2.5646 | 1.9041 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.984 |
|
Br2 |
C1 |
H4 |
117.984 |
| H3 |
C1 |
H4 |
124.033 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
