return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4N4 (2H-Tetrazole, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-294.735404
Energy at 298.15K-294.742647
HF Energy-294.069260
Nuclear repulsion energy222.672054
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3339 3176 0.10      
2 A 3221 3064 0.41      
3 A 3205 3049 4.95      
4 A 3118 2966 8.74      
5 A 1596 1519 13.54      
6 A 1578 1502 11.67      
7 A 1503 1430 1.19      
8 A 1355 1289 6.35      
9 A 1317 1253 21.62      
10 A 1256 1195 0.21      
11 A 1184 1126 0.00      
12 A 1173 1116 10.96      
13 A 1094 1041 10.39      
14 A 1000 951 1.81      
15 A 953 906 1.13      
16 A 902 858 20.83      
17 A 897 854 9.77      
18 A 869 827 18.36      
19 A 687 653 8.63      
20 A 633 602 0.89      
21 A 630 599 12.37      
22 A 320 304 4.34      
23 A 209 199 7.52      
24 A 77 73 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 16057.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15275.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.29879 0.12166 0.08792

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 2.326 -1.247 -0.000
C2 1.446 -0.630 -0.000
N3 0.181 -1.169 0.000
N4 1.500 0.747 0.000
N5 0.157 1.194 -0.000
N6 -0.596 -0.015 -0.000
H7 -2.427 -0.548 -0.895
H8 -2.391 1.011 -0.003
H9 -2.427 -0.542 0.898
C10 -2.076 -0.032 0.000

Atom - Atom Distances (Å)
  H1 C2 N3 N4 N5 N6 H7 H8 H9 C10
H11.07542.14672.15813.26583.17144.88685.23004.88824.5664
C21.07541.37521.37722.23322.13233.97544.17303.97623.5717
N32.14671.37522.32592.36401.39142.82623.37202.82832.5269
N42.15811.37722.32591.41522.23004.23013.89984.22903.6592
N53.26582.23322.36401.41521.42503.24242.55493.24042.5477
N63.17142.13231.39142.23001.42502.10602.06772.10621.4796
H74.88683.97542.82624.23013.24242.10601.79651.79311.0909
H85.23004.17303.37203.89982.55492.06771.79651.79651.0902
H94.88823.97622.82834.22903.24042.10621.79311.79651.0909
C104.56643.57172.52693.65922.54771.47961.09091.09021.0909

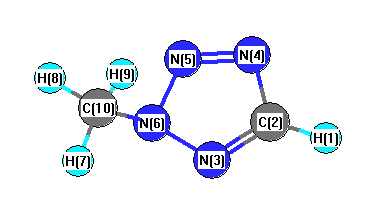
picture of 2H-Tetrazole, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 N3 121.850 H1 C2 N4 122.793
C2 N3 N6 100.838 C2 N4 N5 106.206
N3 C2 N4 115.357 N3 N6 N5 114.135
N3 N6 C10 123.288 N4 N5 N6 103.464
N5 N6 C10 122.578 N6 C10 H7 109.095
N6 C10 H8 106.157 N6 C10 H9 109.110
H7 C10 H8 110.910 H7 C10 H9 110.539
H8 C10 H9 110.905
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability