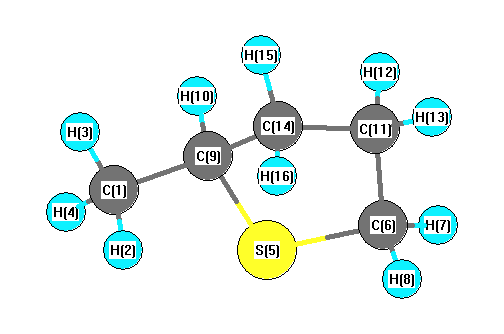
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3018 |
25.85 |
|
|
|
| 2 |
A |
3142 |
3014 |
17.14 |
|
|
|
| 3 |
A |
3127 |
3000 |
14.73 |
|
|
|
| 4 |
A |
3123 |
2996 |
18.78 |
|
|
|
| 5 |
A |
3116 |
2990 |
11.55 |
|
|
|
| 6 |
A |
3083 |
2958 |
24.18 |
|
|
|
| 7 |
A |
3075 |
2950 |
7.10 |
|
|
|
| 8 |
A |
3073 |
2948 |
16.80 |
|
|
|
| 9 |
A |
3063 |
2939 |
19.61 |
|
|
|
| 10 |
A |
3052 |
2928 |
18.89 |
|
|
|
| 11 |
A |
1559 |
1495 |
10.67 |
|
|
|
| 12 |
A |
1547 |
1484 |
6.72 |
|
|
|
| 13 |
A |
1545 |
1482 |
3.14 |
|
|
|
| 14 |
A |
1531 |
1469 |
10.09 |
|
|
|
| 15 |
A |
1527 |
1465 |
4.92 |
|
|
|
| 16 |
A |
1456 |
1397 |
10.48 |
|
|
|
| 17 |
A |
1389 |
1332 |
2.17 |
|
|
|
| 18 |
A |
1382 |
1326 |
0.70 |
|
|
|
| 19 |
A |
1355 |
1300 |
2.79 |
|
|
|
| 20 |
A |
1327 |
1273 |
7.56 |
|
|
|
| 21 |
A |
1316 |
1263 |
17.12 |
|
|
|
| 22 |
A |
1265 |
1213 |
4.96 |
|
|
|
| 23 |
A |
1224 |
1174 |
6.45 |
|
|
|
| 24 |
A |
1166 |
1119 |
0.29 |
|
|
|
| 25 |
A |
1122 |
1077 |
0.98 |
|
|
|
| 26 |
A |
1109 |
1064 |
1.79 |
|
|
|
| 27 |
A |
1058 |
1015 |
8.27 |
|
|
|
| 28 |
A |
1032 |
990 |
0.45 |
|
|
|
| 29 |
A |
1003 |
962 |
5.09 |
|
|
|
| 30 |
A |
967 |
927 |
4.44 |
|
|
|
| 31 |
A |
904 |
867 |
0.78 |
|
|
|
| 32 |
A |
881 |
845 |
3.95 |
|
|
|
| 33 |
A |
852 |
817 |
2.11 |
|
|
|
| 34 |
A |
686 |
658 |
1.79 |
|
|
|
| 35 |
A |
636 |
611 |
2.33 |
|
|
|
| 36 |
A |
538 |
516 |
1.13 |
|
|
|
| 37 |
A |
473 |
454 |
0.10 |
|
|
|
| 38 |
A |
415 |
398 |
0.75 |
|
|
|
| 39 |
A |
293 |
281 |
1.23 |
|
|
|
| 40 |
A |
259 |
248 |
0.18 |
|
|
|
| 41 |
A |
240 |
230 |
0.15 |
|
|
|
| 42 |
A |
92 |
88 |
1.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31570.7 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 30288.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.609 |
|
|
|
| 2 |
H |
0.225 |
|
|
|
| 3 |
H |
0.213 |
|
|
|
| 4 |
H |
0.223 |
|
|
|
| 5 |
S |
0.175 |
|
|
|
| 6 |
C |
-0.588 |
|
|
|
| 7 |
H |
0.242 |
|
|
|
| 8 |
H |
0.239 |
|
|
|
| 9 |
C |
-0.418 |
|
|
|
| 10 |
H |
0.243 |
|
|
|
| 11 |
C |
-0.440 |
|
|
|
| 12 |
H |
0.229 |
|
|
|
| 13 |
H |
0.225 |
|
|
|
| 14 |
C |
-0.411 |
|
|
|
| 15 |
H |
0.223 |
|
|
|
| 16 |
H |
0.226 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.351 |
2.143 |
0.037 |
2.172 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.081 |
-0.330 |
0.951 |
| y |
-0.330 |
-48.041 |
-0.371 |
| z |
0.951 |
-0.371 |
-45.859 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.869 |
-0.330 |
0.951 |
| y |
-0.330 |
-4.071 |
-0.371 |
| z |
0.951 |
-0.371 |
-0.798 |
|
| Polar |
|---|
| 3z2-r2 | -1.596 |
|---|
| x2-y2 | 5.960 |
|---|
| xy | -0.330 |
|---|
| xz | 0.951 |
|---|
| yz | -0.371 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
204.721 |
| (<r2>)1/2 |
14.308 |