return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (1-Butanethiol, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-591.498459
Energy at 298.15K-591.511487
HF Energy-590.912524
Nuclear repulsion energy304.078439
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3170 3016 19.26      
2 A 3161 3007 18.48      
3 A 3160 3006 42.99      
4 A 3152 2999 26.86      
5 A 3148 2995 18.44      
6 A 3112 2960 13.23      
7 A 3101 2950 16.56      
8 A 3084 2934 19.51      
9 A 3082 2931 8.50      
10 A 3075 2926 11.01      
11 A 3068 2918 11.50      
12 A 2780 2645 11.42      
13 A 1609 1530 7.85      
14 A 1602 1524 7.71      
15 A 1601 1523 7.21      
16 A 1599 1521 1.72      
17 A 1583 1506 3.33      
18 A 1575 1498 7.41      
19 A 1494 1421 0.61      
20 A 1492 1419 14.06      
21 A 1440 1370 2.06      
22 A 1422 1353 1.29      
23 A 1396 1328 0.91      
24 A 1377 1310 11.65      
25 A 1340 1275 6.65      
26 A 1295 1232 3.77      
27 A 1222 1163 0.36      
28 A 1189 1131 2.91      
29 A 1133 1078 2.23      
30 A 1068 1016 3.70      
31 A 1045 994 4.07      
32 A 1029 979 4.92      
33 A 960 913 0.22      
34 A 923 878 4.07      
35 A 900 856 0.77      
36 A 837 796 2.72      
37 A 787 748 6.13      
38 A 736 700 1.30      
39 A 470 447 0.14      
40 A 408 388 0.03      
41 A 395 376 0.81      
42 A 270 257 0.16      
43 A 250 237 0.76      
44 A 226 215 0.19      
45 A 209 199 1.95      
46 A 168 160 19.01      
47 A 106 101 0.24      
48 A 68 65 9.03      

Unscaled Zero Point Vibrational Energy (zpe) 36157.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 34396.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14675 0.04977 0.03984

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.003 1.917 0.001
H2 -1.030 2.089 -0.336
H3 0.055 2.138 1.073
H4 0.661 2.605 -0.535
S5 -2.213 -0.415 -0.170
H6 -2.702 -1.477 0.482
C7 -0.496 -0.541 0.489
H8 -0.490 -0.302 1.558
H9 -0.143 -1.564 0.344
C10 0.436 0.451 -0.253
H11 0.377 0.240 -1.329
C12 2.521 -1.109 -0.214
H13 3.576 -1.158 0.077
H14 2.000 -1.946 0.260
H15 2.456 -1.232 -1.301
C16 1.907 0.252 0.205
H17 2.509 1.060 -0.232
H18 1.954 0.366 1.297

Atom - Atom Distances (Å)
  C1 H2 H3 H4 S5 H6 C7 H8 H9 C10 H11 C12 H13 H14 H15 C16 H17 H18
C11.09471.09701.09603.21704.36272.55322.75403.50061.55012.17353.94604.71914.35904.20182.54212.66362.8129
H21.09471.77901.77932.77394.02222.80683.09743.82042.19932.52724.78065.65105.08144.91093.50683.68683.8128
H31.09701.77901.78093.63414.58472.79662.54683.77882.17913.07914.27624.92564.59664.77122.78292.98132.6069
H41.09601.77931.78094.18475.38573.50463.76244.33642.18392.51154.16674.79994.81084.30542.76432.42773.1688
S53.21702.77393.63414.18471.33881.84312.44312.42312.78792.91204.78485.84194.50334.87334.19084.94704.4860
H64.36274.02224.58475.38571.33882.39622.72632.56423.75533.96315.28186.29924.73035.46314.93065.83925.0732
C72.55322.80682.79663.50461.84312.39621.09541.09271.54982.16253.14984.13962.87403.52132.54673.48012.7347
H82.75403.09742.54683.76242.44312.72631.09541.78492.16833.06273.58544.41113.25344.20922.80753.74832.5465
H93.50063.82043.77884.33642.42312.56421.09271.78492.18042.51522.75953.75072.17823.09422.74253.77503.0051
C101.55012.19932.17912.18392.78793.75531.54982.16832.18041.09892.60503.54452.90862.83141.55432.16062.1711
H112.17352.52723.07912.51152.91203.96312.16253.06272.51521.09892.76773.76393.15292.54772.16742.53403.0661
C123.94604.78064.27624.16674.78485.28183.14983.58542.75952.60502.76771.09571.09431.09591.55072.16962.1871
H134.71915.65104.92564.79995.84196.29924.13964.41113.75073.54453.76391.09571.77181.77722.18862.48142.5390
H144.35905.08144.59664.81084.50334.73032.87403.25342.17822.90863.15291.09431.77181.77692.20083.08892.5350
H154.20184.91094.77124.30544.87335.46313.52134.20923.09422.83142.54771.09591.77721.77692.18462.52983.0918
C162.54213.50682.78292.76434.19084.93062.54672.80752.74251.55432.16741.55072.18862.20082.18461.09841.0989
H172.66363.68682.98132.42774.94705.83923.48013.74833.77502.16062.53402.16962.48143.08892.52981.09841.7688
H182.81293.81282.60693.16884.48605.07322.73472.54653.00512.17113.06612.18712.53902.53503.09181.09891.7688

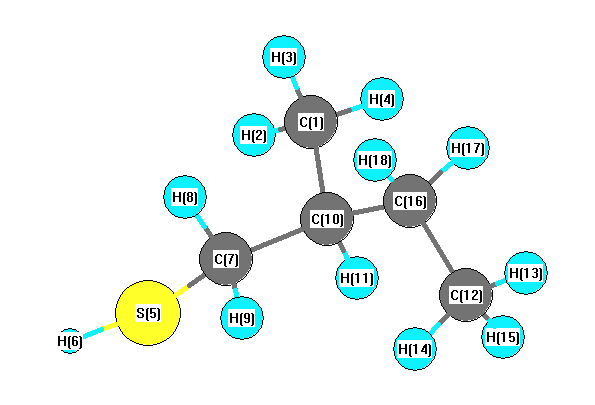
picture of 1-Butanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C10 C7 110.904 C1 C10 H11 109.083
C1 C10 C16 109.946 H2 C1 H3 108.522
H2 C1 H4 108.622 H2 C1 C10 111.352
H3 C1 H4 108.601 H3 C1 C10 109.624
H4 C1 C10 110.059 S5 C7 H8 109.862
S5 C7 H9 108.541 S5 C7 C10 110.209
H6 S5 C7 96.429 C7 C10 H11 108.255
C7 C10 C16 110.260 H8 C7 H9 109.316
H8 C7 C10 108.898 H9 C7 C10 110.003
C10 C16 C12 114.061 C10 C16 H17 107.844
C10 C16 H18 108.616 H11 C10 C16 108.329
C12 C16 H17 108.767 C12 C16 H18 110.096
H13 C12 H14 108.005 H13 C12 H15 108.372
H13 C12 C16 110.399 H14 C12 H15 108.442
H14 C12 C16 111.456 H15 C12 C16 110.076
H17 C16 H18 107.218
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability