return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Butanenitrile, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-248.222815
Energy at 298.15K-248.232448
HF Energy-247.643713
Nuclear repulsion energy217.922067
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3181 3027 16.45      
2 A 3170 3016 25.93      
3 A 3168 3014 22.09      
4 A 3156 3003 22.28      
5 A 3133 2980 5.01      
6 A 3094 2943 7.22      
7 A 3091 2941 18.39      
8 A 3086 2936 10.91      
9 A 3084 2934 1.71      
10 A 2133 2029 3.15      
11 A 1612 1533 4.19      
12 A 1604 1526 9.21      
13 A 1598 1520 9.10      
14 A 1596 1518 5.98      
15 A 1582 1505 0.79      
16 A 1498 1425 5.54      
17 A 1487 1415 6.10      
18 A 1434 1364 0.55      
19 A 1402 1334 0.47      
20 A 1384 1317 0.57      
21 A 1351 1285 0.16      
22 A 1241 1180 1.54      
23 A 1173 1116 4.05      
24 A 1166 1109 4.94      
25 A 1046 995 2.97      
26 A 1032 982 1.05      
27 A 996 947 4.94      
28 A 897 854 0.13      
29 A 842 801 2.22      
30 A 778 740 2.59      
31 A 590 561 0.42      
32 A 555 528 1.24      
33 A 410 390 0.54      
34 A 344 327 0.23      
35 A 294 280 0.23      
36 A 230 219 0.30      
37 A 217 206 1.02      
38 A 180 171 3.46      
39 A 92 88 1.04      

Unscaled Zero Point Vibrational Energy (zpe) 29464.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 28029.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.21392 0.07188 0.05773

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.532 -0.294 0.123
N2 -2.635 -0.686 -0.085
C3 -0.022 1.656 -0.153
H4 -0.161 1.673 -1.239
H5 -0.775 2.301 0.311
H6 0.970 2.047 0.086
C7 2.332 -0.431 0.091
H8 2.451 -0.452 1.180
H9 3.014 -1.170 -0.344
H10 2.624 0.558 -0.274
C11 0.870 -0.767 -0.297
H12 0.743 -0.712 -1.385
H13 0.640 -1.793 0.015
C14 -0.161 0.197 0.369
H15 0.012 0.188 1.454

Atom - Atom Distances (Å)
  C1 N2 C3 H4 H5 H6 C7 H8 H9 H10 C11 H12 H13 C14 H15
C11.18892.48132.75702.71003.42693.86744.12474.65364.26162.48462.76142.64191.47752.0951
N21.18893.50973.60803.54204.52774.97735.24675.67595.40813.51303.61983.45872.66643.1843
C32.48133.50971.09491.09451.09353.15603.51314.15222.86802.58602.77683.51591.55582.1771
H42.75703.60801.09491.78141.78263.52354.14684.35493.15222.81162.55513.77182.18263.0803
H52.71003.54201.09451.78141.77744.14354.32965.18003.86463.53413.77644.34202.19312.5284
H63.42694.52771.09351.78261.77742.82803.10443.83552.25472.84173.13533.85482.18682.4991
C73.86744.97733.15603.52354.14352.82801.09611.09521.09381.54932.18692.17352.58592.7617
H84.12475.24673.51314.14684.32963.10441.09611.77641.77902.18623.09272.53682.81102.5369
H94.65365.67594.15224.35495.18003.83551.09521.77641.77242.18172.53952.48043.52913.7538
H104.26165.40812.86803.15223.86462.25471.09381.77901.77242.19822.52733.08982.88093.1540
C112.48463.51302.58602.81163.53412.84171.54932.18622.18172.19821.09681.09671.56032.1712
H122.76143.61982.77682.55513.77643.13532.18693.09272.53952.52731.09681.77122.17203.0666
H132.64193.45873.51593.77184.34203.85482.17352.53682.48043.08981.09671.77122.17382.5279
C141.47752.66641.55582.18262.19312.18682.58592.81103.52912.88091.56032.17202.17381.0991
H152.09513.18432.17713.08032.52842.49912.76172.53693.75383.15402.17123.06662.52791.0991

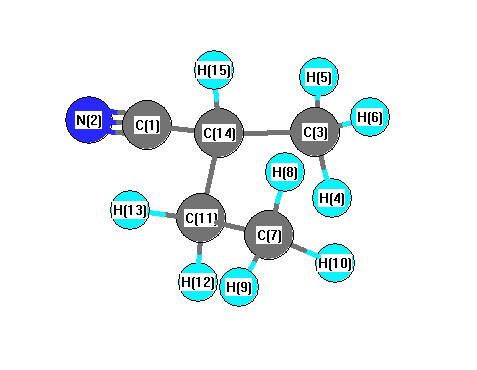
picture of Butanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C14 C3 109.753 C1 C14 C11 109.722
C1 C14 H15 107.906 N2 C1 C14 179.528
C3 C14 C11 112.177 C3 C14 H15 108.965
H4 C3 H5 108.906 H4 C3 H6 109.098
H4 C3 C14 109.631 H5 C3 H6 108.657
H5 C3 C14 110.478 H6 C3 C14 110.040
C7 C11 H12 110.301 C7 C11 H13 109.258
C7 C11 C14 112.519 H8 C7 H9 108.320
H8 C7 H10 108.658 H8 C7 C11 110.288
H9 C7 H10 108.133 H9 C7 C11 109.982
H10 C7 C11 111.376 C11 C14 H15 108.209
H12 C11 H13 107.695 H12 C11 C14 108.395
H13 C11 C14 108.540
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability