Jump to
S1C2
S1C3
Energy calculated at M06-2X/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1585.152372 |
| Energy at 298.15K | |
| HF Energy | -1585.152372 |
| Nuclear repulsion energy | 348.715664 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
502 |
476 |
0.00 |
53.80 |
0.07 |
0.13 |
| 2 |
A1 |
188 |
178 |
0.00 |
3.29 |
0.74 |
0.85 |
| 3 |
B1 |
443 |
420 |
0.00 |
35.45 |
0.75 |
0.86 |
| 4 |
B2 |
329 |
312 |
0.42 |
11.61 |
0.75 |
0.86 |
| 5 |
E |
416 |
394 |
0.00 |
8.13 |
0.75 |
0.86 |
| 5 |
E |
416 |
394 |
0.00 |
8.13 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1147.1 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 1086.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G*
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.439 |
0.313 |
| S2 |
0.000 |
-1.439 |
0.313 |
| S3 |
-1.439 |
0.000 |
-0.313 |
| S4 |
1.439 |
0.000 |
-0.313 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8773 | 2.1288 | 2.1288 |
S2 | 2.8773 | | 2.1288 | 2.1288 | S3 | 2.1288 | 2.1288 | | 2.8773 | S4 | 2.1288 | 2.1288 | 2.8773 | |
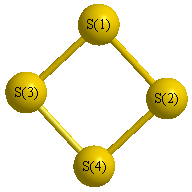 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.482 |
|
S1 |
S3 |
S4 |
47.482 |
| S2 |
S1 |
S3 |
47.482 |
|
S2 |
S4 |
S3 |
47.482 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.328 |
0.000 |
0.000 |
| y |
0.000 |
-50.328 |
0.000 |
| z |
0.000 |
0.000 |
-55.456 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.564 |
0.000 |
0.000 |
| y |
0.000 |
2.564 |
0.000 |
| z |
0.000 |
0.000 |
-5.128 |
|
| Polar |
|---|
| 3z2-r2 | -10.256 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.227 |
0.000 |
0.000 |
| y |
0.000 |
10.227 |
0.000 |
| z |
0.000 |
0.000 |
4.813 |
<r2> (average value of r
2) Å
2
| <r2> |
171.238 |
| (<r2>)1/2 |
13.086 |
Jump to
S1C1
S1C3
Energy calculated at M06-2X/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1585.172407 |
| Energy at 298.15K | |
| HF Energy | -1585.172407 |
| Nuclear repulsion energy | 334.824223 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
708 |
671 |
2.71 |
31.28 |
0.20 |
0.33 |
| 2 |
A1 |
366 |
347 |
7.28 |
47.48 |
0.25 |
0.40 |
| 3 |
A1 |
120 |
114 |
1.39 |
10.81 |
0.51 |
0.68 |
| 4 |
A2 |
224 |
212 |
0.00 |
0.26 |
0.75 |
0.86 |
| 5 |
B2 |
677 |
641 |
173.21 |
2.94 |
0.75 |
0.86 |
| 6 |
B2 |
348 |
330 |
0.11 |
14.62 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1221.7 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 1157.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.049 |
0.915 |
| S2 |
0.000 |
-1.049 |
0.915 |
| S3 |
0.000 |
1.571 |
-0.915 |
| S4 |
0.000 |
-1.571 |
-0.915 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.0989 | 1.9026 | 3.1962 |
S2 | 2.0989 | | 3.1962 | 1.9026 | S3 | 1.9026 | 3.1962 | | 3.1423 | S4 | 3.1962 | 1.9026 | 3.1423 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
105.914 |
|
S1 |
S3 |
S4 |
74.086 |
| S2 |
S1 |
S3 |
105.914 |
|
S2 |
S4 |
S3 |
74.086 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.098 |
|
|
|
| 2 |
S |
0.098 |
|
|
|
| 3 |
S |
-0.098 |
|
|
|
| 4 |
S |
-0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.629 |
1.629 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.836 |
0.000 |
0.000 |
| y |
0.000 |
-54.632 |
0.000 |
| z |
0.000 |
0.000 |
-50.318 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.639 |
0.000 |
0.000 |
| y |
0.000 |
-3.555 |
0.000 |
| z |
0.000 |
0.000 |
2.916 |
|
| Polar |
|---|
| 3z2-r2 | 5.833 |
|---|
| x2-y2 | 2.796 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.913 |
0.000 |
0.000 |
| y |
0.000 |
17.530 |
0.000 |
| z |
0.000 |
0.000 |
10.586 |
<r2> (average value of r
2) Å
2
| <r2> |
200.446 |
| (<r2>)1/2 |
14.158 |
Jump to
S1C1
S1C2
Energy calculated at M06-2X/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1585.168735 |
| Energy at 298.15K | |
| HF Energy | -1585.168735 |
| Nuclear repulsion energy | 336.127491 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
768 |
727 |
0.00 |
36.22 |
0.30 |
0.46 |
| 2 |
Ag |
319 |
302 |
0.00 |
48.61 |
0.29 |
0.45 |
| 3 |
Au |
256 |
242 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
B1u |
702 |
665 |
200.24 |
0.00 |
0.00 |
0.00 |
| 5 |
B2u |
118i |
112i |
19.93 |
0.00 |
0.00 |
0.00 |
| 6 |
B3g |
350 |
332 |
0.00 |
17.91 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1138.3 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 1078.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G*
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.941 |
1.272 |
| S2 |
0.000 |
0.941 |
-1.272 |
| S3 |
0.000 |
-0.941 |
1.272 |
| S4 |
0.000 |
-0.941 |
-1.272 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5442 | 1.8814 | 3.1643 |
S2 | 2.5442 | | 3.1643 | 1.8814 | S3 | 1.8814 | 3.1643 | | 2.5442 | S4 | 3.1643 | 1.8814 | 2.5442 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.239 |
0.000 |
0.000 |
| y |
0.000 |
-49.785 |
0.000 |
| z |
0.000 |
0.000 |
-54.011 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.341 |
0.000 |
0.000 |
| y |
0.000 |
3.339 |
0.000 |
| z |
0.000 |
0.000 |
-2.999 |
|
| Polar |
|---|
| 3z2-r2 | -5.997 |
|---|
| x2-y2 | -2.453 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.853 |
0.000 |
0.000 |
| y |
0.000 |
11.186 |
0.000 |
| z |
0.000 |
0.000 |
16.742 |
<r2> (average value of r
2) Å
2
| <r2> |
192.688 |
| (<r2>)1/2 |
13.881 |
