return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,3-Hexadiene, (E)-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.236632
Energy at 298.15K-232.246451
HF Energy-231.696875
Nuclear repulsion energy211.518369
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3259 3100 15.32      
2 A' 3172 3018 4.85      
3 A' 3171 3016 14.74      
4 A' 3157 3004 31.45      
5 A' 3152 2999 7.51      
6 A' 3141 2988 16.44      
7 A' 3086 2936 13.68      
8 A' 3066 2917 18.96      
9 A' 1700 1617 5.28      
10 A' 1658 1577 2.68      
11 A' 1608 1530 5.44      
12 A' 1581 1504 2.68      
13 A' 1524 1450 0.76      
14 A' 1491 1418 6.00      
15 A' 1426 1357 7.64      
16 A' 1393 1325 4.04      
17 A' 1367 1301 0.16      
18 A' 1352 1286 2.88      
19 A' 1216 1157 5.12      
20 A' 1105 1051 0.35      
21 A' 1068 1016 0.77      
22 A' 974 926 1.92      
23 A' 880 837 1.49      
24 A' 579 551 2.48      
25 A' 483 460 0.14      
26 A' 322 307 0.21      
27 A' 156 149 0.37      
28 A" 3164 3010 26.92      
29 A" 3095 2944 13.68      
30 A" 1602 1524 5.73      
31 A" 1360 1294 0.00      
32 A" 1171 1114 0.85      
33 A" 1046 995 53.07      
34 A" 997 949 5.77      
35 A" 937 891 44.11      
36 A" 881 838 0.70      
37 A" 794 755 7.49      
38 A" 633 602 1.76      
39 A" 308 293 1.73      
40 A" 226 215 1.45      
41 A" 145 138 0.69      
42 A" 97 92 0.12      

Unscaled Zero Point Vibrational Energy (zpe) 31771.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30224.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.34336 0.05106 0.04520

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.003 3.986 0.000
H2 1.456 2.846 0.000
C3 0.375 2.967 0.000
H4 -1.534 2.065 0.000
C5 -0.455 1.904 0.000
H6 1.079 0.344 0.000
C7 0.000 0.503 0.000
H8 -1.918 -0.362 0.000
C9 -0.842 -0.552 0.000
H10 -0.889 -2.506 0.881
H11 -0.889 -2.506 -0.881
C12 -0.445 -2.022 0.000
H13 1.287 -3.351 0.000
H14 1.548 -1.835 -0.889
H15 1.548 -1.835 0.889
C16 1.084 -2.274 0.000

Atom - Atom Distances (Å)
  H1 H2 C3 H4 C5 H6 C7 H8 C9 H10 H11 C12 H13 H14 H15 C16
H11.84791.08532.46072.13193.79803.48394.75334.61656.61266.61266.02477.44876.08856.08856.3535
H21.84791.08783.09072.13042.53002.75884.65544.10225.90945.90945.22536.19864.76554.76555.1335
C31.08531.08782.11201.34872.71572.49304.04233.72395.68625.68625.05576.38335.02275.02275.2891
H42.46073.09072.11201.09123.12832.18932.45632.70664.69934.69934.22896.10615.04945.04945.0676
C52.13192.13041.34871.09122.18741.47352.69722.48674.51844.51843.92585.53604.33404.33404.4528
H63.79802.53002.71573.12832.18741.09023.07862.11953.57393.57392.81363.70062.40002.40002.6183
C73.48392.75882.49302.18931.47351.09022.10371.34973.25873.25872.56304.06262.94142.94142.9810
H84.75334.65544.04232.45632.69723.07862.10371.09262.53642.53642.21964.38273.86973.86973.5597
C94.61654.10223.72392.70662.48672.11951.34971.09262.14392.14391.52233.51662.85472.85472.5839
H106.61265.90945.68624.69934.51843.57393.25872.53642.14391.76161.09932.49543.08602.52792.1736
H116.61265.90945.68624.69934.51843.57393.25872.53642.14391.76161.09932.49542.52793.08602.1736
C126.02475.22535.05574.22893.92582.81362.56302.21961.52231.09931.09932.18302.18982.18981.5494
H137.44876.19866.38336.10615.53603.70064.06264.38273.51662.49542.49542.18301.77631.77631.0953
H146.08854.76555.02275.04944.33402.40002.94143.86972.85473.08602.52792.18981.77631.77871.0948
H156.08854.76555.02275.04944.33402.40002.94143.86972.85472.52793.08602.18981.77631.77871.0948
C166.35355.13355.28915.06764.45282.61832.98103.55972.58392.17362.17361.54941.09531.09481.0948

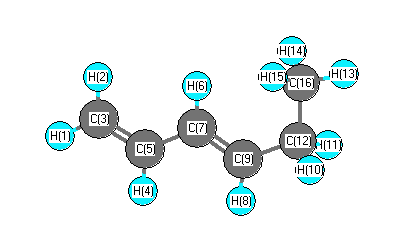
picture of 1,3-Hexadiene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C3 H2 116.501 H1 C3 C5 121.925
H2 C3 C5 121.573 C3 C5 H4 119.534
C3 C5 C7 124.042 H4 C5 C7 116.424
C5 C7 H6 116.333 C5 C7 C9 123.423
H6 C7 C9 120.244 C7 C9 H8 118.563
C7 C9 C12 126.253 H8 C9 C12 115.184
C9 C12 H10 108.651 C9 C12 H11 108.651
C9 C12 C16 114.527 H10 C12 H11 106.493
H10 C12 C16 109.108 H11 C12 C16 109.108
C12 C16 H13 110.072 C12 C16 H14 110.633
C12 C16 H15 110.633 H13 C16 H14 108.396
H13 C16 H15 108.396 H14 C16 H15 108.644
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability