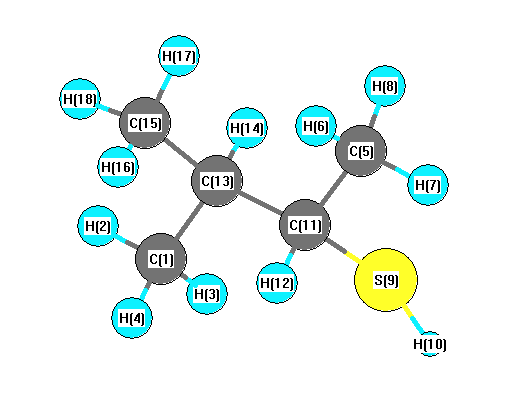
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3017 |
19.10 |
|
|
|
| 2 |
A |
3140 |
3013 |
29.77 |
|
|
|
| 3 |
A |
3136 |
3008 |
21.89 |
|
|
|
| 4 |
A |
3127 |
3000 |
54.02 |
|
|
|
| 5 |
A |
3126 |
2999 |
10.58 |
|
|
|
| 6 |
A |
3118 |
2992 |
0.72 |
|
|
|
| 7 |
A |
3076 |
2951 |
16.92 |
|
|
|
| 8 |
A |
3059 |
2934 |
15.31 |
|
|
|
| 9 |
A |
3056 |
2932 |
6.52 |
|
|
|
| 10 |
A |
3052 |
2928 |
11.68 |
|
|
|
| 11 |
A |
3050 |
2926 |
13.30 |
|
|
|
| 12 |
A |
2700 |
2590 |
19.21 |
|
|
|
| 13 |
A |
1581 |
1516 |
4.66 |
|
|
|
| 14 |
A |
1563 |
1500 |
17.23 |
|
|
|
| 15 |
A |
1560 |
1497 |
22.43 |
|
|
|
| 16 |
A |
1549 |
1486 |
2.97 |
|
|
|
| 17 |
A |
1548 |
1486 |
4.50 |
|
|
|
| 18 |
A |
1541 |
1478 |
2.67 |
|
|
|
| 19 |
A |
1467 |
1407 |
8.23 |
|
|
|
| 20 |
A |
1457 |
1397 |
26.36 |
|
|
|
| 21 |
A |
1445 |
1386 |
10.01 |
|
|
|
| 22 |
A |
1393 |
1336 |
0.62 |
|
|
|
| 23 |
A |
1370 |
1315 |
0.43 |
|
|
|
| 24 |
A |
1351 |
1296 |
1.50 |
|
|
|
| 25 |
A |
1283 |
1231 |
23.43 |
|
|
|
| 26 |
A |
1220 |
1170 |
4.59 |
|
|
|
| 27 |
A |
1204 |
1155 |
8.90 |
|
|
|
| 28 |
A |
1160 |
1113 |
2.08 |
|
|
|
| 29 |
A |
1103 |
1059 |
6.74 |
|
|
|
| 30 |
A |
1057 |
1014 |
12.04 |
|
|
|
| 31 |
A |
1019 |
977 |
6.20 |
|
|
|
| 32 |
A |
992 |
952 |
0.09 |
|
|
|
| 33 |
A |
966 |
927 |
2.24 |
|
|
|
| 34 |
A |
929 |
892 |
2.51 |
|
|
|
| 35 |
A |
916 |
879 |
4.78 |
|
|
|
| 36 |
A |
795 |
763 |
3.15 |
|
|
|
| 37 |
A |
697 |
669 |
2.28 |
|
|
|
| 38 |
A |
483 |
463 |
0.45 |
|
|
|
| 39 |
A |
415 |
398 |
0.38 |
|
|
|
| 40 |
A |
378 |
363 |
0.19 |
|
|
|
| 41 |
A |
372 |
357 |
0.39 |
|
|
|
| 42 |
A |
340 |
326 |
1.56 |
|
|
|
| 43 |
A |
266 |
255 |
4.34 |
|
|
|
| 44 |
A |
245 |
235 |
2.36 |
|
|
|
| 45 |
A |
236 |
227 |
0.29 |
|
|
|
| 46 |
A |
214 |
205 |
6.10 |
|
|
|
| 47 |
A |
188 |
180 |
11.45 |
|
|
|
| 48 |
A |
64 |
61 |
1.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35574.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 34130.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.595 |
|
|
|
| 2 |
H |
0.209 |
|
|
|
| 3 |
H |
0.236 |
|
|
|
| 4 |
H |
0.208 |
|
|
|
| 5 |
C |
-0.609 |
|
|
|
| 6 |
H |
0.220 |
|
|
|
| 7 |
H |
0.212 |
|
|
|
| 8 |
H |
0.228 |
|
|
|
| 9 |
S |
-0.076 |
|
|
|
| 10 |
H |
0.129 |
|
|
|
| 11 |
C |
-0.390 |
|
|
|
| 12 |
H |
0.240 |
|
|
|
| 13 |
C |
-0.288 |
|
|
|
| 14 |
H |
0.230 |
|
|
|
| 15 |
C |
-0.590 |
|
|
|
| 16 |
H |
0.207 |
|
|
|
| 17 |
H |
0.213 |
|
|
|
| 18 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.974 |
1.434 |
0.648 |
1.851 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.340 |
2.756 |
1.795 |
| y |
2.756 |
-47.821 |
0.683 |
| z |
1.795 |
0.683 |
-47.859 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.500 |
2.756 |
1.795 |
| y |
2.756 |
-0.721 |
0.683 |
| z |
1.795 |
0.683 |
-0.779 |
|
| Polar |
|---|
| 3z2-r2 | -1.558 |
|---|
| x2-y2 | 1.481 |
|---|
| xy | 2.756 |
|---|
| xz | 1.795 |
|---|
| yz | 0.683 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
249.896 |
| (<r2>)1/2 |
15.808 |