Jump to
S1C2
Vibrational Frequencies calculated at MP2/3-21G*
Geometric Data calculated at MP2/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -191.519520 |
| Energy at 298.15K | -191.524362 |
| HF Energy | -191.114683 |
| Nuclear repulsion energy | 147.550709 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3300 |
3139 |
0.29 |
|
|
|
| 2 |
A1 |
3283 |
3123 |
5.74 |
|
|
|
| 3 |
A1 |
3258 |
3100 |
1.09 |
|
|
|
| 4 |
A1 |
1544 |
1469 |
0.03 |
|
|
|
| 5 |
A1 |
1427 |
1358 |
1.41 |
|
|
|
| 6 |
A1 |
1179 |
1122 |
0.42 |
|
|
|
| 7 |
A1 |
1115 |
1061 |
13.68 |
|
|
|
| 8 |
A1 |
1014 |
965 |
6.75 |
|
|
|
| 9 |
A1 |
895 |
851 |
0.20 |
|
|
|
| 10 |
A2 |
850 |
808 |
0.00 |
|
|
|
| 11 |
A2 |
811 |
771 |
0.00 |
|
|
|
| 12 |
A2 |
539 |
512 |
0.00 |
|
|
|
| 13 |
B1 |
893 |
850 |
11.75 |
|
|
|
| 14 |
B1 |
725 |
690 |
19.53 |
|
|
|
| 15 |
B1 |
677 |
644 |
106.85 |
|
|
|
| 16 |
B1 |
525 |
500 |
13.32 |
|
|
|
| 17 |
B2 |
3710 |
3529 |
9874.47 |
|
|
|
| 18 |
B2 |
3259 |
3100 |
190.82 |
|
|
|
| 19 |
B2 |
3254 |
3095 |
96.21 |
|
|
|
| 20 |
B2 |
1401 |
1333 |
72.05 |
|
|
|
| 21 |
B2 |
1349 |
1284 |
4.04 |
|
|
|
| 22 |
B2 |
1210 |
1151 |
12.33 |
|
|
|
| 23 |
B2 |
945 |
899 |
0.85 |
|
|
|
| 24 |
B2 |
865 |
823 |
8.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19013.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 18087.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.228 |
| C2 |
0.000 |
1.134 |
0.404 |
| C3 |
0.000 |
0.688 |
-1.019 |
| C4 |
0.000 |
-0.688 |
-1.019 |
| C5 |
0.000 |
-1.134 |
0.404 |
| H6 |
0.000 |
0.000 |
2.308 |
| H7 |
0.000 |
2.166 |
0.729 |
| H8 |
0.000 |
1.345 |
-1.877 |
| H9 |
0.000 |
-1.345 |
-1.877 |
| H10 |
0.000 |
-2.166 |
0.729 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4017 | 2.3495 | 2.3495 | 1.4017 | 1.0797 | 2.2227 | 3.3838 | 3.3838 | 2.2227 |
C2 | 1.4017 | | 1.4905 | 2.3111 | 2.2675 | 2.2160 | 1.0822 | 2.2904 | 3.3684 | 3.3157 | C3 | 2.3495 | 1.4905 | | 1.3759 | 2.3111 | 3.3967 | 2.2887 | 1.0811 | 2.2069 | 3.3464 | C4 | 2.3495 | 2.3111 | 1.3759 | | 1.4905 | 3.3967 | 3.3464 | 2.2069 | 1.0811 | 2.2887 | C5 | 1.4017 | 2.2675 | 2.3111 | 1.4905 | | 2.2160 | 3.3157 | 3.3684 | 2.2904 | 1.0822 | H6 | 1.0797 | 2.2160 | 3.3967 | 3.3967 | 2.2160 | | 2.6803 | 4.3955 | 4.3955 | 2.6803 | H7 | 2.2227 | 1.0822 | 2.2887 | 3.3464 | 3.3157 | 2.6803 | | 2.7321 | 4.3725 | 4.3319 | H8 | 3.3838 | 2.2904 | 1.0811 | 2.2069 | 3.3684 | 4.3955 | 2.7321 | | 2.6904 | 4.3725 | H9 | 3.3838 | 3.3684 | 2.2069 | 1.0811 | 2.2904 | 4.3955 | 4.3725 | 2.6904 | | 2.7321 | H10 | 2.2227 | 3.3157 | 3.3464 | 2.2887 | 1.0822 | 2.6803 | 4.3319 | 4.3725 | 2.7321 | |
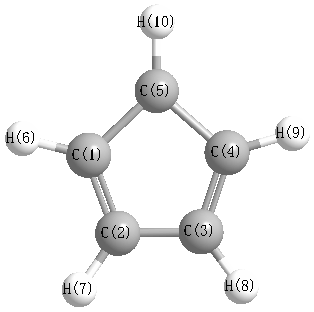 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.614 |
|
C1 |
C2 |
H7 |
126.490 |
| C1 |
C5 |
C4 |
108.614 |
|
C1 |
C5 |
H10 |
126.490 |
| C2 |
C1 |
C5 |
107.963 |
|
C2 |
C1 |
H6 |
126.019 |
| C2 |
C3 |
C4 |
107.404 |
|
C2 |
C3 |
H8 |
125.153 |
| C3 |
C2 |
H7 |
124.896 |
|
C3 |
C4 |
C5 |
107.404 |
| C3 |
C4 |
H9 |
127.443 |
|
C4 |
C3 |
H8 |
127.443 |
| C4 |
C5 |
H10 |
124.896 |
|
C5 |
C1 |
H6 |
126.019 |
| C5 |
C4 |
H9 |
125.153 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
