Jump to
S2C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -76.488792 |
| Energy at 298.15K | -76.488672 |
| HF Energy | -76.333548 |
| Nuclear repulsion energy | 23.609441 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3177 |
3022 |
45.27 |
|
|
|
| 2 |
A1 |
1642 |
1562 |
80.05 |
|
|
|
| 3 |
A1 |
1330 |
1265 |
19.11 |
|
|
|
| 4 |
B1 |
843 |
802 |
133.11 |
|
|
|
| 5 |
B2 |
3258 |
3099 |
19.87 |
|
|
|
| 6 |
B2 |
510 |
485 |
1.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5379.6 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 5117.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.485 |
| C2 |
0.000 |
0.000 |
0.833 |
| H3 |
0.000 |
0.933 |
-1.044 |
| H4 |
0.000 |
-0.933 |
-1.044 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3174 | 1.0880 | 1.0880 |
C2 | 1.3174 | | 2.0963 | 2.0963 | H3 | 1.0880 | 2.0963 | | 1.8660 | H4 | 1.0880 | 2.0963 | 1.8660 | |
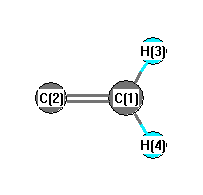 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
120.964 |
|
C2 |
C1 |
H4 |
120.964 |
| H3 |
C1 |
H4 |
118.073 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -76.427902 |
| Energy at 298.15K | -76.428037 |
| HF Energy | -76.311926 |
| Nuclear repulsion energy | 23.558304 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3134 |
2981 |
1.69 |
|
|
|
| 2 |
A1 |
1582 |
1505 |
0.00 |
|
|
|
| 3 |
A1 |
1490 |
1417 |
7.57 |
|
|
|
| 4 |
B1 |
975 |
927 |
53.13 |
|
|
|
| 5 |
B2 |
3206 |
3050 |
7.00 |
|
|
|
| 6 |
B2 |
1095 |
1042 |
4.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5740.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 5461.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.484 |
| C2 |
0.000 |
0.000 |
0.837 |
| H3 |
0.000 |
0.925 |
-1.058 |
| H4 |
0.000 |
-0.925 |
-1.058 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3204 | 1.0887 | 1.0887 |
C2 | 1.3204 | | 2.1085 | 2.1085 | H3 | 1.0887 | 2.1085 | | 1.8497 | H4 | 1.0887 | 2.1085 | 1.8497 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
