return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2 (benzyl radical)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2B1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-268.180990
Energy at 298.15K-268.188086
HF Energy-267.639866
Nuclear repulsion energy260.277532
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3238 3080 12.41      
2 A1 3213 3057 1.11      
3 A1 3196 3040 6.88      
4 A1 3188 3032 6.57      
5 A1 1648 1568 2.01      
6 A1 1566 1489 1.41      
7 A1 1548 1473 10.54      
8 A1 1303 1240 1.82      
9 A1 1224 1165 1.13      
10 A1 1078 1026 1.34      
11 A1 1005 956 0.01      
12 A1 853 811 0.42      
13 A1 567 539 0.45      
14 A2 1031 981 0.00      
15 A2 895 851 0.00      
16 A2 499 475 0.00      
17 A2 420 399 0.00      
18 B1 1020 970 1.08      
19 B1 941 895 1.70      
20 B1 779 741 103.14      
21 B1 720 685 5.74      
22 B1 664 632 39.21      
23 B1 499 475 17.23      
24 B1 216 205 1.71      
25 B2 3290 3129 6.57      
26 B2 3221 3064 25.08      
27 B2 3197 3041 2.17      
28 B2 1630 1550 2.71      
29 B2 1535 1460 8.93      
30 B2 1426 1357 1.10      
31 B2 1290 1227 0.07      
32 B2 1183 1125 3.35      
33 B2 1170 1113 0.02      
34 B2 1010 961 0.83      
35 B2 674 641 0.00      
36 B2 377 359 0.17      

Unscaled Zero Point Vibrational Energy (zpe) 25654.2 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 24404.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.18547 0.09038 0.06077

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.986
C2 0.000 1.216 0.247
C3 0.000 1.208 -1.129
C4 0.000 0.000 -1.833
C5 0.000 -1.208 -1.129
C6 0.000 -1.216 0.247
C7 0.000 0.000 2.403
H8 0.000 2.158 0.790
H9 0.000 2.147 -1.674
H10 0.000 0.000 -2.918
H11 0.000 -2.147 -1.674
H12 0.000 -2.158 0.790
H13 0.000 0.928 2.965
H14 0.000 -0.928 2.965

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.42242.43552.81902.43551.42241.41702.16683.41853.90413.41852.16682.18562.1856
C21.42241.37642.40952.78702.43112.47471.08752.13533.39093.87313.41692.73263.4610
C32.43551.37641.39812.41572.78703.73272.14141.08622.15873.39933.87444.10344.6174
C42.81902.40951.39811.39812.40954.23593.39672.15341.08522.15343.39674.88674.8867
C52.43552.78702.41571.39811.37643.73273.87443.39932.15871.08622.14144.61744.1034
C61.42242.43112.78702.40951.37642.47473.41693.87313.39092.13531.08753.46102.7326
C71.41702.47473.73274.23593.73272.47472.69404.60775.32114.60772.69401.08471.0847
H82.16681.08752.14143.39673.87443.41692.69402.46394.29054.96064.31582.49853.7751
H93.41852.13531.08622.15343.39933.87314.60772.46392.48204.29504.96064.79635.5654
H103.90413.39092.15871.08522.15873.39095.32114.29052.48202.48204.29055.95575.9557
H113.41853.87313.39932.15341.08622.13534.60774.96064.29502.48202.46395.56544.7963
H122.16683.41693.87443.39672.14141.08752.69404.31584.96064.29052.46393.77512.4985
H132.18562.73264.10344.88674.61743.46101.08472.49854.79635.95575.56543.77511.8557
H142.18563.46104.61744.88674.10342.73261.08473.77515.56545.95574.79632.49851.8557

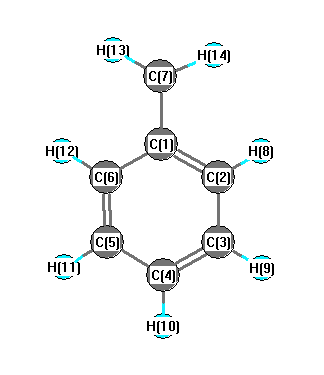
picture of benzyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.963 C1 C2 H8 118.777
C1 C6 C5 120.963 C1 C6 H12 118.777
C1 C7 H13 121.199 C1 C7 H14 121.199
C2 C1 C6 117.433 C2 C1 C7 121.283
C2 C3 C4 120.558 C2 C3 H9 119.786
C3 C2 H8 120.260 C3 C4 C5 119.526
C3 C4 H10 120.237 C4 C3 H9 119.656
C4 C5 C6 120.558 C4 C5 H11 119.656
C5 C1 C7 150.269 C5 C4 H10 120.237
C5 C6 H12 120.260 C6 C5 H11 119.786
H13 C7 H14 117.601
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability