return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6S3 (Carbonotrithioic acid, dimethyl ester)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-1304.088009
Energy at 298.15K-1304.094265
HF Energy-1303.435431
Nuclear repulsion energy430.130115
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3197 3042 2.73      
2 A 3196 3041 0.03      
3 A 3188 3033 1.87      
4 A 3187 3032 3.05      
5 A 3097 2946 3.75      
6 A 3096 2946 5.48      
7 A 1567 1490 1.68      
8 A 1564 1488 33.14      
9 A 1555 1479 24.17      
10 A 1555 1479 0.85      
11 A 1463 1391 0.48      
12 A 1456 1385 0.08      
13 A 1133 1078 118.58      
14 A 1059 1007 9.31      
15 A 1050 998 14.48      
16 A 1049 998 26.12      
17 A 1048 997 0.00      
18 A 832 792 171.54      
19 A 730 694 2.25      
20 A 715 680 1.02      
21 A 513 488 2.97      
22 A 498 473 1.18      
23 A 363 345 6.00      
24 A 317 301 2.10      
25 A 259 247 5.62      
26 A 197 188 1.12      
27 A 192 183 0.01      
28 A 189 180 0.00      
29 A 90 86 5.75      
30 A 58 55 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 19205.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 18270.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.10467 0.04482 0.03176

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.000 0.020 -0.000
S2 0.000 1.663 0.000
S3 1.437 -1.008 0.000
S4 -1.437 -1.008 -0.000
C5 2.767 0.231 -0.000
C6 -2.767 0.231 0.000
H7 2.703 0.858 0.891
H8 -2.704 0.857 0.892
H9 3.704 -0.334 -0.000
H10 2.703 0.857 -0.892
H11 -2.704 0.858 -0.891
H12 -3.704 -0.334 -0.000

Atom - Atom Distances (Å)
  C1 S2 S3 S4 C5 C6 H7 H8 H9 H10 H11 H12
C11.64281.76751.76742.77492.77482.96732.96743.72082.96702.96733.7208
S21.64283.03343.03343.11523.11562.95792.95884.20772.95812.95834.2081
S31.76753.03342.87471.81764.38312.42494.62852.36482.42484.62875.1854
S41.76743.03342.87474.38301.81754.62862.42495.18534.62812.42482.3648
C52.77493.11521.81764.38305.53361.09165.57791.09421.09165.57776.4954
C62.77483.11564.38311.81755.53365.57761.09166.49545.57751.09161.0942
H72.96732.95792.42494.62861.09165.57765.40691.79331.78305.69306.5779
H82.96742.95884.62852.42495.57791.09165.40696.57805.69331.78311.7933
H93.72084.20772.36485.18531.09426.49541.79336.57801.79346.57807.4079
H102.96702.95812.42484.62811.09165.57751.78305.69331.79345.40666.5776
H112.96732.95834.62872.42485.57771.09165.69301.78316.57805.40661.7933
H123.72084.20815.18542.36486.49541.09426.57791.79337.40796.57761.7933

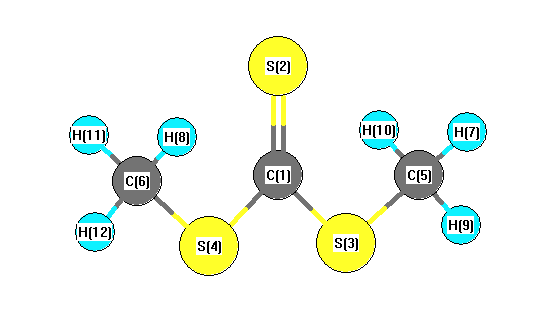
picture of Carbonotrithioic acid, dimethyl ester state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S3 C5 101.419 C1 S4 C6 101.429
S2 C1 S3 125.581 S2 C1 S4 125.589
S3 C1 S4 108.830 S3 C5 H7 110.424
S3 C5 H9 105.928 S3 C5 H10 110.416
S4 C6 H8 110.430 S4 C6 H11 110.430
S4 C6 H12 105.930 H7 C5 H9 110.254
H7 C5 H10 109.511 H8 C6 H11 109.509
H8 C6 H12 110.249 H9 C5 H10 110.263
H11 C6 H12 110.248
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability