return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2NH2 (diaminomethane)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-149.712262
Energy at 298.15K-149.720272
HF Energy-149.400750
Counterpoise corrected energy-149.712262
CP Energy at 298.15K-149.720272
Counterpoise optimized geometry corrected energy 
CP opt. Energy at 298.15K 
Nuclear repulsion energy81.783368
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3428 3250 5.23      
2 A1 3118 2956 35.37      
3 A1 1766 1675 32.67      
4 A1 1570 1489 0.79      
5 A1 1100 1043 57.48      
6 A1 783 743 5.46      
7 A1 445 422 5.76      
8 A2 3534 3350 0.00      
9 A2 1448 1372 0.00      
10 A2 1104 1046 0.00      
11 A2 299 284 0.00      
12 B1 3534 3350 2.93      
13 B1 3173 3008 26.78      
14 B1 1404 1331 0.29      
15 B1 847 803 0.09      
16 B1 440 417 120.75      
17 B2 3430 3251 0.47      
18 B2 1754 1662 0.33      
19 B2 1436 1361 6.75      
20 B2 1050 995 13.10      
21 B2 696 660 563.27      

Unscaled Zero Point Vibrational Energy (zpe) 18179.0 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 17233.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
1.16446 0.28836 0.26280

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
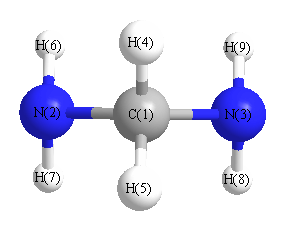
C1 0.000 0.000 0.560
N2 0.000 1.286 -0.191
N3 0.000 -1.286 -0.191
H4 0.885 0.000 1.200
H5 -0.885 0.000 1.200
H6 0.836 1.384 -0.773
H7 -0.836 1.384 -0.773
H8 -0.836 -1.384 -0.773
H9 0.836 -1.384 -0.773

Atom - Atom Distances (Å)
  C1 N2 N3 H4 H5 H6 H7 H8 H9
C11.48931.48931.09241.09242.09562.09562.09562.0956
N21.48932.57272.09102.09101.02331.02332.85832.8583
N31.48932.57272.09102.09102.85832.85831.02331.0233
H41.09242.09102.09101.77042.41062.96152.96152.4106
H51.09242.09102.09101.77042.96152.41062.41062.9615
H62.09561.02332.85832.41062.96151.67163.23422.7687
H72.09561.02332.85832.96152.41061.67162.76873.2342
H82.09562.85831.02332.96152.41063.23422.76871.6716
H92.09562.85831.02332.41062.96152.76873.23421.6716

picture of diaminomethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 111.684 C1 N2 H7 111.684
C1 N3 H8 111.684 C1 N3 H9 111.684
N2 C1 N3 119.468 N2 C1 H4 107.177
N2 C1 H5 107.177 N3 C1 H4 107.177
N3 C1 H5 107.177 H4 C1 H5 108.263
H6 N2 H7 109.533 H8 N3 H9 109.533
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability