return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5 (phenyl)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-229.234444
Energy at 298.15K-229.239599
HF Energy-228.781435
Nuclear repulsion energy195.730273
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3247 3088 0.58      
2 A1 3234 3076 9.29      
3 A1 3207 3051 0.08      
4 A1 1701 1618 0.80      
5 A1 1569 1493 20.37      
6 A1 1277 1214 0.00      
7 A1 1118 1064 9.75      
8 A1 1060 1009 0.47      
9 A1 1026 976 0.24      
10 A1 683 650 1.06      
11 A2 1093 1040 0.00      
12 A2 957 910 0.00      
13 A2 472 449 0.00      
14 B1 1087 1034 0.12      
15 B1 1016 967 1.97      
16 B1 819 779 70.60      
17 B1 775 737 51.78      
18 B1 496 472 5.42      
19 B2 3242 3084 7.56      
20 B2 3216 3059 6.53      
21 B2 1748 1663 5.79      
22 B2 1552 1476 10.00      
23 B2 1400 1332 0.03      
24 B2 1293 1230 0.34      
25 B2 1181 1123 2.44      
26 B2 1104 1050 2.76      
27 B2 666 633 0.33      

Unscaled Zero Point Vibrational Energy (zpe) 20118.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 19138.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.21214 0.19022 0.10029

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.397
C2 0.000 1.214 0.761
C3 0.000 -1.214 0.761
C4 0.000 1.202 -0.626
C5 0.000 -1.202 -0.626
C6 0.000 0.000 -1.314
H7 0.000 2.148 1.313
H8 0.000 -2.148 1.313
H9 0.000 2.141 -1.172
H10 0.000 -2.141 -1.172
H11 0.000 0.000 -2.400

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11
C11.37081.37082.35372.35372.71162.14952.14953.34453.34453.7969
C21.37082.42871.38752.78652.40471.08433.40712.14383.87263.3861
C31.37082.42872.78651.38752.40473.40711.08433.87262.14383.3861
C42.35371.38752.78652.40441.38522.15743.87081.08613.38782.1425
C52.35372.78651.38752.40441.38523.87082.15743.38781.08612.1425
C62.71162.40472.40471.38521.38523.39343.39342.14612.14611.0853
H72.14951.08433.40712.15743.87083.39344.29572.48464.95694.2890
H82.14953.40711.08433.87082.15743.39344.29574.95692.48464.2890
H93.34452.14383.87261.08613.38782.14612.48464.95694.28272.4684
H103.34453.87262.14383.38781.08612.14614.95692.48464.28272.4684
H113.79693.38613.38612.14252.14251.08534.28904.28902.46842.4684

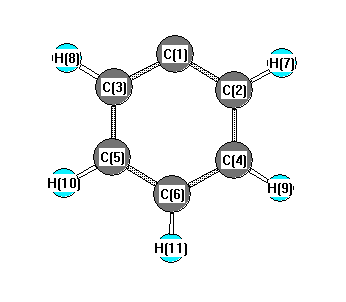
picture of phenyl state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 117.145 C1 C2 H7 121.774
C1 C3 C5 117.145 C1 C3 H8 121.774
C2 C1 C3 124.709 C2 C4 C6 120.285
C2 C4 H9 119.652 C3 C5 C6 120.285
C3 C5 H10 119.652 C4 C2 H7 121.081
C4 C6 C5 120.430 C4 C6 H11 119.785
C5 C3 H8 121.081 C5 C6 H11 119.785
C6 C4 H9 120.063 C6 C5 H10 120.063
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability