Jump to
S1C2
Vibrational Frequencies calculated at MP2/3-21G*
Geometric Data calculated at MP2/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -233.423376 |
| Energy at 298.15K | -233.436905 |
| HF Energy | -232.873575 |
| Nuclear repulsion energy | 247.536382 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3169 |
3015 |
57.88 |
|
|
|
| 2 |
A' |
3150 |
2997 |
41.25 |
|
|
|
| 3 |
A' |
3142 |
2989 |
18.70 |
|
|
|
| 4 |
A' |
3125 |
2973 |
11.59 |
|
|
|
| 5 |
A' |
3112 |
2961 |
4.85 |
|
|
|
| 6 |
A' |
3097 |
2946 |
4.49 |
|
|
|
| 7 |
A' |
3072 |
2922 |
19.62 |
|
|
|
| 8 |
A' |
3069 |
2919 |
20.65 |
|
|
|
| 9 |
A' |
1600 |
1522 |
12.27 |
|
|
|
| 10 |
A' |
1598 |
1521 |
0.73 |
|
|
|
| 11 |
A' |
1578 |
1502 |
1.03 |
|
|
|
| 12 |
A' |
1490 |
1417 |
9.51 |
|
|
|
| 13 |
A' |
1487 |
1415 |
0.65 |
|
|
|
| 14 |
A' |
1421 |
1352 |
0.11 |
|
|
|
| 15 |
A' |
1405 |
1337 |
5.74 |
|
|
|
| 16 |
A' |
1319 |
1255 |
5.82 |
|
|
|
| 17 |
A' |
1283 |
1221 |
1.32 |
|
|
|
| 18 |
A' |
1232 |
1172 |
0.50 |
|
|
|
| 19 |
A' |
1127 |
1072 |
0.16 |
|
|
|
| 20 |
A' |
1064 |
1012 |
0.82 |
|
|
|
| 21 |
A' |
1008 |
959 |
3.06 |
|
|
|
| 22 |
A' |
978 |
930 |
0.48 |
|
|
|
| 23 |
A' |
863 |
821 |
1.39 |
|
|
|
| 24 |
A' |
795 |
756 |
1.39 |
|
|
|
| 25 |
A' |
655 |
623 |
1.84 |
|
|
|
| 26 |
A' |
442 |
420 |
0.16 |
|
|
|
| 27 |
A' |
333 |
316 |
0.16 |
|
|
|
| 28 |
A' |
126 |
120 |
0.02 |
|
|
|
| 29 |
A" |
3163 |
3009 |
13.97 |
|
|
|
| 30 |
A" |
3141 |
2988 |
40.90 |
|
|
|
| 31 |
A" |
3141 |
2988 |
3.97 |
|
|
|
| 32 |
A" |
3094 |
2944 |
39.85 |
|
|
|
| 33 |
A" |
1598 |
1520 |
1.94 |
|
|
|
| 34 |
A" |
1597 |
1520 |
1.94 |
|
|
|
| 35 |
A" |
1561 |
1485 |
3.44 |
|
|
|
| 36 |
A" |
1334 |
1269 |
0.16 |
|
|
|
| 37 |
A" |
1320 |
1256 |
0.00 |
|
|
|
| 38 |
A" |
1267 |
1205 |
0.12 |
|
|
|
| 39 |
A" |
1182 |
1124 |
0.95 |
|
|
|
| 40 |
A" |
1170 |
1113 |
0.16 |
|
|
|
| 41 |
A" |
1045 |
994 |
5.05 |
|
|
|
| 42 |
A" |
890 |
846 |
0.21 |
|
|
|
| 43 |
A" |
879 |
837 |
0.04 |
|
|
|
| 44 |
A" |
826 |
786 |
0.24 |
|
|
|
| 45 |
A" |
384 |
365 |
0.01 |
|
|
|
| 46 |
A" |
273 |
260 |
0.36 |
|
|
|
| 47 |
A" |
254 |
241 |
0.00 |
|
|
|
| 48 |
A" |
239 |
227 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37549.6 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35721.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.265 |
0.090 |
1.102 |
| C2 |
0.265 |
0.090 |
-1.102 |
| C3 |
-0.679 |
0.709 |
0.000 |
| C4 |
0.880 |
-0.861 |
0.000 |
| C5 |
-0.894 |
2.225 |
0.000 |
| C6 |
0.265 |
-2.268 |
0.000 |
| H7 |
0.997 |
0.825 |
1.458 |
| H8 |
-0.212 |
-0.391 |
1.963 |
| H9 |
0.997 |
0.825 |
-1.458 |
| H10 |
-0.212 |
-0.391 |
-1.963 |
| H11 |
-1.645 |
0.190 |
0.000 |
| H12 |
1.973 |
-0.926 |
0.000 |
| H13 |
0.074 |
2.742 |
0.000 |
| H14 |
-0.831 |
-2.209 |
0.000 |
| H15 |
-1.454 |
2.542 |
-0.890 |
| H16 |
-1.454 |
2.542 |
0.890 |
| H17 |
0.578 |
-2.828 |
-0.890 |
| H18 |
0.578 |
-2.828 |
0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.2030 | 1.5773 | 1.5791 | 2.6681 | 2.6024 | 1.0970 | 1.0953 | 2.7614 | 3.1380 | 2.2075 | 2.2720 | 2.8786 | 2.7747 | 3.5962 | 3.0023 | 3.5465 | 2.9420 |
C2 | 2.2030 | | 1.5773 | 1.5791 | 2.6681 | 2.6024 | 2.7614 | 3.1380 | 1.0970 | 1.0953 | 2.2075 | 2.2720 | 2.8786 | 2.7747 | 3.0023 | 3.5962 | 2.9420 | 3.5465 | C3 | 1.5773 | 1.5773 | | 2.2116 | 1.5320 | 3.1228 | 2.2239 | 2.2972 | 2.2239 | 2.2972 | 1.0968 | 3.1153 | 2.1687 | 2.9216 | 2.1802 | 2.1802 | 3.8576 | 3.8576 | C4 | 1.5791 | 1.5791 | 2.2116 | | 3.5594 | 1.5360 | 2.2317 | 2.2945 | 2.2317 | 2.2945 | 2.7347 | 1.0953 | 3.6917 | 2.1783 | 4.2205 | 4.2205 | 2.1805 | 2.1805 | C5 | 2.6681 | 2.6681 | 1.5320 | 3.5594 | | 4.6407 | 2.7676 | 3.3406 | 2.7676 | 3.3406 | 2.1698 | 4.2606 | 1.0976 | 4.4350 | 1.0975 | 1.0975 | 5.3383 | 5.3383 | C6 | 2.6024 | 2.6024 | 3.1228 | 1.5360 | 4.6407 | | 3.4972 | 2.7577 | 3.4972 | 2.7577 | 3.1130 | 2.1722 | 5.0141 | 1.0977 | 5.1849 | 5.1849 | 1.0973 | 1.0973 | H7 | 1.0970 | 2.7614 | 2.2239 | 2.2317 | 2.7676 | 3.4972 | | 1.7874 | 2.9152 | 3.8259 | 3.0836 | 2.4790 | 2.5787 | 3.8306 | 3.8026 | 3.0452 | 4.3630 | 3.7209 | H8 | 1.0953 | 3.1380 | 2.2972 | 2.2945 | 3.3406 | 2.7577 | 1.7874 | | 3.8259 | 3.9251 | 2.4984 | 2.9856 | 3.7078 | 2.7462 | 4.2749 | 3.3604 | 3.8347 | 2.7778 | H9 | 2.7614 | 1.0970 | 2.2239 | 2.2317 | 2.7676 | 3.4972 | 2.9152 | 3.8259 | | 1.7874 | 3.0836 | 2.4790 | 2.5787 | 3.8306 | 3.0452 | 3.8026 | 3.7209 | 4.3630 | H10 | 3.1380 | 1.0953 | 2.2972 | 2.2945 | 3.3406 | 2.7577 | 3.8259 | 3.9251 | 1.7874 | | 2.4984 | 2.9856 | 3.7078 | 2.7462 | 3.3604 | 4.2749 | 2.7778 | 3.8347 | H11 | 2.2075 | 2.2075 | 1.0968 | 2.7347 | 2.1698 | 3.1130 | 3.0836 | 2.4984 | 3.0836 | 2.4984 | | 3.7865 | 3.0776 | 2.5333 | 2.5222 | 2.5222 | 3.8528 | 3.8528 | H12 | 2.2720 | 2.2720 | 3.1153 | 1.0953 | 4.2606 | 2.1722 | 2.4790 | 2.9856 | 2.4790 | 2.9856 | 3.7865 | | 4.1307 | 3.0837 | 4.9559 | 4.9559 | 2.5210 | 2.5210 | H13 | 2.8786 | 2.8786 | 2.1687 | 3.6917 | 1.0976 | 5.0141 | 2.5787 | 3.7078 | 2.5787 | 3.7078 | 3.0776 | 4.1307 | | 5.0334 | 1.7792 | 1.7792 | 5.6635 | 5.6635 | H14 | 2.7747 | 2.7747 | 2.9216 | 2.1783 | 4.4350 | 1.0977 | 3.8306 | 2.7462 | 3.8306 | 2.7462 | 2.5333 | 3.0837 | 5.0334 | | 4.8737 | 4.8737 | 1.7781 | 1.7781 | H15 | 3.5962 | 3.0023 | 2.1802 | 4.2205 | 1.0975 | 5.1849 | 3.8026 | 4.2749 | 3.0452 | 3.3604 | 2.5222 | 4.9559 | 1.7792 | 4.8737 | | 1.7791 | 5.7416 | 6.0112 | H16 | 3.0023 | 3.5962 | 2.1802 | 4.2205 | 1.0975 | 5.1849 | 3.0452 | 3.3604 | 3.8026 | 4.2749 | 2.5222 | 4.9559 | 1.7792 | 4.8737 | 1.7791 | | 6.0112 | 5.7416 | H17 | 3.5465 | 2.9420 | 3.8576 | 2.1805 | 5.3383 | 1.0973 | 4.3630 | 3.8347 | 3.7209 | 2.7778 | 3.8528 | 2.5210 | 5.6635 | 1.7781 | 5.7416 | 6.0112 | | 1.7805 | H18 | 2.9420 | 3.5465 | 3.8576 | 2.1805 | 5.3383 | 1.0973 | 3.7209 | 2.7778 | 4.3630 | 3.8347 | 3.8528 | 2.5210 | 5.6635 | 1.7781 | 6.0112 | 5.7416 | 1.7805 | |
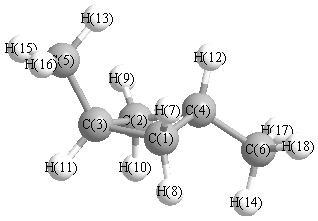 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.592 |
|
C1 |
C3 |
C5 |
118.204 |
| C1 |
C3 |
H11 |
109.984 |
|
C1 |
C4 |
C2 |
88.459 |
| C1 |
C4 |
C6 |
113.308 |
|
C1 |
C4 |
H12 |
115.117 |
| C2 |
C3 |
C5 |
118.204 |
|
C2 |
C3 |
H11 |
109.984 |
| C2 |
C4 |
C6 |
113.308 |
|
C2 |
C4 |
H12 |
115.117 |
| C3 |
C1 |
C4 |
88.964 |
|
C3 |
C1 |
H7 |
111.260 |
| C3 |
C1 |
H8 |
117.394 |
|
C3 |
C2 |
C4 |
88.964 |
| C3 |
C2 |
H9 |
111.260 |
|
C3 |
C2 |
H10 |
117.394 |
| C3 |
C5 |
H13 |
110.019 |
|
C3 |
C5 |
H15 |
110.933 |
| C3 |
C5 |
H16 |
110.933 |
|
C4 |
C1 |
H7 |
111.747 |
| C4 |
C1 |
H8 |
117.015 |
|
C4 |
C2 |
H9 |
111.747 |
| C4 |
C2 |
H10 |
117.015 |
|
C4 |
C6 |
H14 |
110.503 |
| C4 |
C6 |
H17 |
110.696 |
|
C4 |
C6 |
H18 |
110.696 |
| C5 |
C3 |
H11 |
110.155 |
|
C6 |
C4 |
H12 |
110.156 |
| H7 |
C1 |
H8 |
109.237 |
|
H9 |
C2 |
H10 |
109.237 |
| H13 |
C5 |
H15 |
108.289 |
|
H13 |
C5 |
H16 |
108.289 |
| H14 |
C6 |
H17 |
108.206 |
|
H14 |
C6 |
H18 |
108.206 |
| H15 |
C5 |
H16 |
108.290 |
|
H17 |
C6 |
H18 |
108.448 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
