Jump to
S1C2
S2C1
S2C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -38.761890 |
| Energy at 298.15K | -38.761712 |
| HF Energy | -38.708953 |
| Nuclear repulsion energy | 6.150513 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
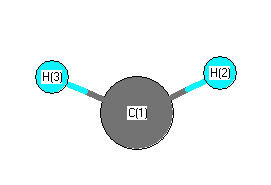
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.113 |
| H2 |
0.000 |
0.981 |
-0.338 |
| H3 |
0.000 |
-0.981 |
-0.338 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0798 | 1.0798 |
H2 | 1.0798 | | 1.9629 | H3 | 1.0798 | 1.9629 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
130.717 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -38.748761 |
| Energy at 298.15K | |
| HF Energy | -38.697435 |
| Nuclear repulsion energy | 6.186351 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
3235 |
3078 |
0.00 |
|
|
|
| 2 |
Σu |
3575 |
3401 |
29.28 |
|
|
|
| 3 |
Πu |
1153i |
1097i |
127.06 |
|
|
|
| 3 |
Πu |
1153i |
1097i |
127.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 2252.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 2143.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.000 |
| H2 |
0.000 |
0.000 |
1.069 |
| H3 |
0.000 |
0.000 |
-1.069 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0692 | 1.0692 |
H2 | 1.0692 | | 2.1385 | H3 | 1.0692 | 2.1385 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -38.717108 |
| Energy at 298.15K | -38.716947 |
| HF Energy | -38.651549 |
| Nuclear repulsion energy | 5.979649 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.173 |
| H2 |
0.000 |
0.878 |
-0.520 |
| H3 |
0.000 |
-0.878 |
-0.520 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.1183 | 1.1183 |
H2 | 1.1183 | | 1.7551 | H3 | 1.1183 | 1.7551 | |
More geometry information
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -38.717108 |
| Energy at 298.15K | -38.716947 |
| HF Energy | -38.651549 |
| Nuclear repulsion energy | 5.979649 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Geometric Data calculated at MP2/3-21G*
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
