return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Pentalene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-305.321035
Energy at 298.15K-305.326359
HF Energy-304.617963
Nuclear repulsion energy310.375259
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3267 3107 0.00      
2 Ag 3243 3085 0.00      
3 Ag 3233 3075 0.00      
4 Ag 1603 1525 0.00      
5 Ag 1520 1446 0.00      
6 Ag 1371 1304 0.00      
7 Ag 1303 1239 0.00      
8 Ag 1172 1115 0.00      
9 Ag 1150 1094 0.00      
10 Ag 899 855 0.00      
11 Ag 833 793 0.00      
12 Ag 748 712 0.00      
13 Ag 646 614 0.00      
14 Au 853 811 7.74      
15 Au 790 752 32.06      
16 Au 686 652 110.23      
17 Au 483 460 2.36      
18 Au 290 276 2.62      
19 Au 96 91 7.63      
20 Bg 869 827 0.00      
21 Bg 816 776 0.00      
22 Bg 707 673 0.00      
23 Bg 610 580 0.00      
24 Bg 530 504 0.00      
25 Bu 3266 3107 23.28      
26 Bu 3243 3085 9.19      
27 Bu 3234 3076 6.78      
28 Bu 1596 1518 10.81      
29 Bu 1490 1417 4.92      
30 Bu 1365 1298 12.42      
31 Bu 1291 1228 11.34      
32 Bu 1155 1099 20.00      
33 Bu 1030 980 15.45      
34 Bu 896 852 0.97      
35 Bu 877 834 5.32      
36 Bu 437 416 3.36      

Unscaled Zero Point Vibrational Energy (zpe) 23797.6 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 22638.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.16864 0.07060 0.04977

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.361 1.111 0.000
C2 2.192 -0.173 0.000
C3 1.361 -1.260 0.000
C4 -1.361 -1.111 0.000
C5 -2.192 0.173 0.000
C6 -1.361 1.260 0.000
C7 0.046 0.745 0.000
C8 -0.046 -0.745 0.000
H9 1.788 2.108 0.000
H10 3.276 -0.186 0.000
H11 1.656 -2.302 0.000
H12 -1.788 -2.108 0.000
H13 -3.276 0.186 0.000
H14 -1.656 2.302 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.52902.37073.51253.67452.72521.36442.32861.08492.31263.42514.50214.72773.2431
C21.52901.36913.67454.39803.83082.33432.31032.31591.08362.19594.42555.47974.5749
C32.37071.36912.72523.83083.70902.39791.49783.39492.19611.08273.26044.85654.6676
C43.51253.67452.72521.52902.37072.32861.36444.50214.72773.24311.08492.31263.4251
C53.67454.39803.83081.52901.36912.31032.33434.42555.47974.57492.31591.08362.1959
C62.72523.83083.70902.37071.36911.49782.39793.26044.85654.66763.39492.19611.0827
C71.36442.33432.39792.32862.31031.49781.49352.21133.36133.44633.39153.36862.3062
C82.32862.31031.49781.36442.33432.39791.49353.39153.36862.30622.21133.36133.4463
H91.08492.31593.39494.50214.42553.26042.21133.39152.73384.41165.52765.41613.4490
H102.31261.08362.19614.72775.47974.85653.36133.36862.73382.66515.41616.56215.5234
H113.42512.19591.08273.24314.57494.66763.44632.30624.41162.66513.44905.52345.6711
H124.50214.42553.26041.08492.31593.39493.39152.21135.52765.41613.44902.73384.4116
H134.72775.47974.85652.31261.08362.19613.36863.36135.41616.56215.52342.73382.6651
H143.24314.57494.66763.42512.19591.08272.30623.44633.44905.52345.67114.41162.6651

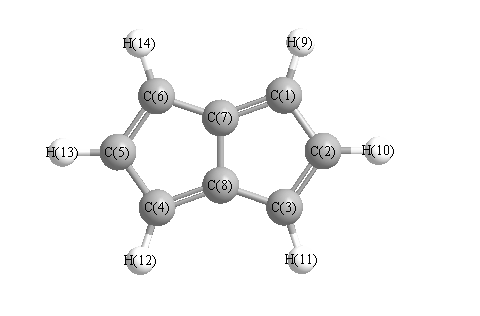
picture of Pentalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 109.645 C1 C2 H10 123.642
C1 C7 C6 144.371 C1 C7 C8 109.053
C2 C1 C7 107.426 C2 C1 H9 123.860
C2 C3 C8 107.300 C2 C3 H11 126.773
C3 C2 H10 126.713 C3 C8 C4 144.371
C3 C8 C7 106.576 C4 C5 C6 109.645
C4 C5 H13 123.642 C4 C8 C7 109.053
C5 C4 C8 107.426 C5 C4 H12 123.860
C5 C6 C7 107.300 C5 C6 H14 126.773
C6 C5 H13 126.713 C6 C7 C8 106.576
C7 C1 H9 128.714 C7 C6 H14 125.927
C8 C3 H11 125.927 C8 C4 H12 128.714
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability