Jump to
S1C2
Energy calculated at wB97X-D/3-21G*
| | hartrees |
|---|
| Energy at 0K | -416.263597 |
| Energy at 298.15K | -416.267794 |
| HF Energy | -416.263597 |
| Nuclear repulsion energy | 61.638301 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3629 |
3440 |
16.39 |
|
|
|
| 2 |
A' |
2301 |
2181 |
125.50 |
|
|
|
| 3 |
A' |
1214 |
1151 |
21.98 |
|
|
|
| 4 |
A' |
1142 |
1082 |
82.40 |
|
|
|
| 5 |
A' |
938 |
889 |
48.97 |
|
|
|
| 6 |
A' |
847 |
803 |
173.26 |
|
|
|
| 7 |
A" |
2291 |
2171 |
192.43 |
|
|
|
| 8 |
A" |
928 |
880 |
29.00 |
|
|
|
| 9 |
A" |
483 |
458 |
160.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6886.1 cm
-1
Scaled (by 0.9478) Zero Point Vibrational Energy (zpe) 6526.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
-0.105 |
-0.562 |
0.000 |
| O2 |
-0.105 |
1.088 |
0.000 |
| H3 |
0.761 |
1.546 |
0.000 |
| H4 |
0.829 |
-0.912 |
1.024 |
| H5 |
0.829 |
-0.912 |
-1.024 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.6496 | 2.2785 | 1.4294 | 1.4294 |
O2 | 1.6496 | | 0.9799 | 2.4331 | 2.4331 | H3 | 2.2785 | 0.9799 | | 2.6633 | 2.6633 | H4 | 1.4294 | 2.4331 | 2.6633 | | 2.0482 | H5 | 1.4294 | 2.4331 | 2.6633 | 2.0482 | |
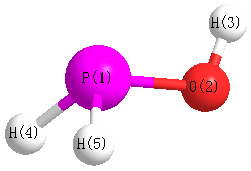 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
117.849 |
|
O2 |
P1 |
H4 |
104.184 |
| O2 |
P1 |
H5 |
104.184 |
|
H4 |
P1 |
H5 |
91.525 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.305 |
|
|
|
| 2 |
O |
-0.633 |
|
|
|
| 3 |
H |
0.381 |
|
|
|
| 4 |
H |
-0.027 |
|
|
|
| 5 |
H |
-0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.469 |
0.358 |
0.000 |
2.495 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.188 |
1.882 |
0.000 |
| y |
1.882 |
-19.088 |
0.000 |
| z |
0.000 |
0.000 |
-19.848 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.719 |
1.882 |
0.000 |
| y |
1.882 |
0.930 |
0.000 |
| z |
0.000 |
0.000 |
-0.210 |
|
| Polar |
|---|
| 3z2-r2 | -0.421 |
|---|
| x2-y2 | -1.100 |
|---|
| xy | 1.882 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.891 |
-0.325 |
0.000 |
| y |
-0.325 |
3.414 |
0.000 |
| z |
0.000 |
0.000 |
3.204 |
<r2> (average value of r
2) Å
2
| <r2> |
34.748 |
| (<r2>)1/2 |
5.895 |
Jump to
S1C1
Energy calculated at wB97X-D/3-21G*
| | hartrees |
|---|
| Energy at 0K | -416.263423 |
| Energy at 298.15K | -416.267414 |
| HF Energy | -416.263423 |
| Nuclear repulsion energy | 61.494434 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3660 |
3469 |
55.79 |
|
|
|
| 2 |
A' |
2392 |
2267 |
93.80 |
|
|
|
| 3 |
A' |
1212 |
1149 |
34.12 |
|
|
|
| 4 |
A' |
1160 |
1099 |
144.60 |
|
|
|
| 5 |
A' |
952 |
903 |
16.10 |
|
|
|
| 6 |
A' |
833 |
789 |
122.38 |
|
|
|
| 7 |
A" |
2381 |
2256 |
138.75 |
|
|
|
| 8 |
A" |
965 |
914 |
2.27 |
|
|
|
| 9 |
A" |
274 |
259 |
119.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6913.7 cm
-1
Scaled (by 0.9478) Zero Point Vibrational Energy (zpe) 6552.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
0.039 |
-0.568 |
0.000 |
| O2 |
0.039 |
1.082 |
0.000 |
| H3 |
0.936 |
1.470 |
0.000 |
| H4 |
-0.918 |
-0.804 |
1.018 |
| H5 |
-0.918 |
-0.804 |
-1.018 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.6491 | 2.2257 | 1.4170 | 1.4170 |
O2 | 1.6491 | | 0.9768 | 2.3469 | 2.3469 | H3 | 2.2257 | 0.9768 | | 3.1051 | 3.1051 | H4 | 1.4170 | 2.3469 | 3.1051 | | 2.0359 | H5 | 1.4170 | 2.3469 | 3.1051 | 2.0359 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
113.413 |
|
O2 |
P1 |
H4 |
99.613 |
| O2 |
P1 |
H5 |
99.613 |
|
H4 |
P1 |
H5 |
91.841 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.260 |
|
|
|
| 2 |
O |
-0.630 |
|
|
|
| 3 |
H |
0.384 |
|
|
|
| 4 |
H |
-0.007 |
|
|
|
| 5 |
H |
-0.007 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.990 |
0.110 |
0.000 |
0.996 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.174 |
4.281 |
0.000 |
| y |
4.281 |
-19.776 |
0.000 |
| z |
0.000 |
0.000 |
-19.691 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.560 |
4.281 |
0.000 |
| y |
4.281 |
-0.344 |
0.000 |
| z |
0.000 |
0.000 |
-0.216 |
|
| Polar |
|---|
| 3z2-r2 | -0.433 |
|---|
| x2-y2 | 0.602 |
|---|
| xy | 4.281 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.101 |
0.877 |
0.000 |
| y |
0.877 |
3.332 |
0.000 |
| z |
0.000 |
0.000 |
3.206 |
<r2> (average value of r
2) Å
2
| <r2> |
34.745 |
| (<r2>)1/2 |
5.894 |
