Jump to
S1C2
S2C1
S2C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -130.168969 |
| Energy at 298.15K | -130.168654 |
| HF Energy | -129.950448 |
| Nuclear repulsion energy | 47.309260 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3339 |
3165 |
7.02 |
|
|
|
| 2 |
A' |
2130 |
2019 |
13.12 |
|
|
|
| 3 |
A' |
1070 |
1015 |
27.70 |
|
|
|
| 4 |
A' |
860 |
815 |
2.36 |
|
|
|
| 5 |
A' |
496 |
470 |
3.15 |
|
|
|
| 6 |
A" |
580 |
550 |
3.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4237.4 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 4017.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.311 |
-1.222 |
0.000 |
| C2 |
0.000 |
0.118 |
0.000 |
| N3 |
0.229 |
1.251 |
0.000 |
| H4 |
0.265 |
-2.131 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3755 | 2.5309 | 1.0765 |
C2 | 1.3755 | | 1.1557 | 2.2648 | N3 | 2.5309 | 1.1557 | | 3.3822 | H4 | 1.0765 | 2.2648 | 3.3822 | |
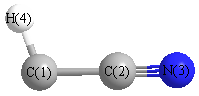 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
178.345 |
|
C2 |
C1 |
H4 |
134.568 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -130.163803 |
| Energy at 298.15K | |
| HF Energy | -129.954965 |
| Nuclear repulsion energy | 47.538138 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3460 |
3280 |
54.02 |
|
|
|
| 2 |
Σ |
1550 |
1469 |
31.54 |
|
|
|
| 3 |
Σ |
1215 |
1152 |
32.87 |
|
|
|
| 4 |
Π |
573 |
543 |
1.30 |
|
|
|
| 4 |
Π |
573 |
543 |
1.30 |
|
|
|
| 5 |
Π |
413i |
392i |
57.32 |
|
|
|
| 5 |
Π |
413i |
392i |
57.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3271.8 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 3101.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.211 |
| C2 |
0.000 |
0.000 |
0.093 |
| N3 |
0.000 |
0.000 |
1.284 |
| H4 |
0.000 |
0.000 |
-2.277 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3040 | 2.4950 | 1.0655 |
C2 | 1.3040 | | 1.1910 | 2.3695 | N3 | 2.4950 | 1.1910 | | 3.5605 | H4 | 1.0655 | 2.3695 | 3.5605 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.013 |
|
|
|
| 2 |
C |
0.091 |
|
|
|
| 3 |
N |
-0.433 |
|
|
|
| 4 |
H |
0.329 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.322 |
3.322 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.318 |
0.000 |
0.000 |
| y |
0.000 |
-17.318 |
0.000 |
| z |
0.000 |
0.000 |
-16.080 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.619 |
0.000 |
0.000 |
| y |
0.000 |
-0.619 |
0.000 |
| z |
0.000 |
0.000 |
1.238 |
|
| Polar |
|---|
| 3z2-r2 | 2.476 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.235 |
0.000 |
0.000 |
| y |
0.000 |
1.235 |
0.000 |
| z |
0.000 |
0.000 |
5.398 |
<r2> (average value of r
2) Å
2
| <r2> |
36.134 |
| (<r2>)1/2 |
6.011 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -130.151732 |
| Energy at 298.15K | -130.151265 |
| HF Energy | -129.877064 |
| Nuclear repulsion energy | 46.130098 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3005 |
2849 |
27.78 |
|
|
|
| 2 |
A' |
2248 |
2131 |
13.79 |
|
|
|
| 3 |
A' |
1111 |
1053 |
85.62 |
|
|
|
| 4 |
A' |
884 |
838 |
1.65 |
|
|
|
| 5 |
A' |
429 |
407 |
18.71 |
|
|
|
| 6 |
A" |
436 |
413 |
10.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4056.7 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 3845.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.003 |
-1.318 |
0.000 |
| C2 |
0.000 |
0.104 |
0.000 |
| N3 |
-0.152 |
1.285 |
0.000 |
| H4 |
1.041 |
-1.707 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4215 | 2.6070 | 1.1082 |
C2 | 1.4215 | | 1.1906 | 2.0888 | N3 | 2.6070 | 1.1906 | | 3.2207 | H4 | 1.1082 | 2.0888 | 3.2207 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -130.151732 |
| Energy at 298.15K | -130.151265 |
| HF Energy | -129.877064 |
| Nuclear repulsion energy | 46.130098 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
