return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H4N2 (Pyridazine)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-261.730393
Energy at 298.15K-261.736116
HF Energy-261.149068
Nuclear repulsion energy203.511746
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3238 3080 1.95      
2 A1 3212 3055 3.64      
3 A1 1575 1498 10.69      
4 A1 1432 1362 6.14      
5 A1 1229 1169 0.65      
6 A1 1120 1066 1.71      
7 A1 1030 980 0.86      
8 A1 794 755 0.96      
9 A1 685 652 1.69      
10 A2 987 939 0.00      
11 A2 950 904 0.00      
12 A2 751 715 0.00      
13 A2 365 347 0.00      
14 B1 982 934 0.26      
15 B1 767 730 53.10      
16 B1 380 362 8.75      
17 B2 3223 3066 15.88      
18 B2 3206 3050 4.64      
19 B2 1538 1463 6.81      
20 B2 1448 1377 16.39      
21 B2 1293 1230 1.70      
22 B2 1092 1039 3.40      
23 B2 1024 975 1.35      
24 B2 632 602 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 16476.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15673.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.19885 0.19200 0.09768

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.696 1.195
C2 0.000 -0.696 1.195
C3 0.000 -1.348 -0.056
C4 0.000 1.348 -0.056
H5 0.000 1.270 2.116
H6 0.000 -1.270 2.116
H7 0.000 -2.432 -0.116
H8 0.000 2.432 -0.116
N9 0.000 0.717 -1.262
N10 0.000 -0.717 -1.262

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 N9 N10
C11.39132.39641.41141.08512.17093.39092.17532.45702.8340
C21.39131.41142.39642.17091.08512.17533.39092.83402.4570
C32.39641.41142.69573.40202.17341.08533.77992.39071.3606
C41.41142.39642.69572.17343.40203.77991.08531.36062.3907
H51.08512.17093.40202.17342.54104.32252.51533.42243.9186
H62.17091.08512.17343.40202.54102.51534.32253.91863.4224
H73.39092.17531.08533.77994.32252.51534.86313.35042.0625
H82.17533.39093.77991.08532.51534.32254.86312.06253.3504
N92.45702.83402.39071.36063.42243.91863.35042.06251.4335
N102.83402.45701.36062.39073.91863.42242.06253.35041.4335

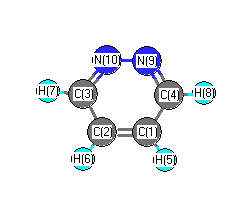
picture of Pyridazine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 117.524 C1 C2 H6 121.988
C1 C4 H8 120.656 C1 C4 N9 124.840
C2 C1 C4 117.524 C2 C1 H5 121.988
C2 C3 H7 120.656 C2 C3 N10 124.840
C3 C2 H6 120.488 C3 N10 N9 117.636
C4 C1 H5 120.488 C4 N9 N10 117.636
H7 C3 N10 114.504 H8 C4 N9 114.504
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability