return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5NO (3-Methylisoxazole)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-282.636465
Energy at 298.15K-282.642843
HF Energy-282.030335
Nuclear repulsion energy217.913221
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3323 3162 0.33      
2 A 3297 3136 0.09      
3 A 3177 3022 4.40      
4 A 3162 3008 10.31      
5 A 3094 2943 9.82      
6 A 1598 1520 9.16      
7 A 1589 1512 8.87      
8 A 1534 1459 12.96      
9 A 1496 1423 5.13      
10 A 1427 1358 18.80      
11 A 1352 1286 12.67      
12 A 1295 1232 1.64      
13 A 1154 1098 9.97      
14 A 1132 1077 2.88      
15 A 1050 999 11.84      
16 A 1014 965 5.18      
17 A 973 925 1.88      
18 A 917 873 0.35      
19 A 857 815 0.33      
20 A 850 808 9.71      
21 A 775 738 57.37      
22 A 657 625 0.28      
23 A 637 606 5.67      
24 A 584 556 5.14      
25 A 336 320 5.18      
26 A 264 251 4.91      
27 A 134 127 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 18838.5 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 17921.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.29015 0.11282 0.08250

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.162 0.033 -0.000
H2 -2.543 0.544 -0.890
H3 -2.525 -0.999 -0.000
H4 -2.543 0.544 0.890
C5 1.499 0.625 0.000
O6 1.449 -0.778 -0.000
N7 0.004 -1.157 0.000
C8 -0.652 0.021 -0.000
C9 0.238 1.153 0.000
H10 -0.039 2.195 0.000
H11 2.489 1.054 -0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 O6 N7 C8 C9 H10 H11
C11.09491.09391.09493.70863.70022.47121.50932.64803.02964.7608
H21.09491.78101.78084.13984.29783.18942.15402.98263.12855.1348
H31.09391.78101.78104.33953.97942.53382.13193.50174.04715.4171
H41.09491.78081.78104.13994.29783.18932.15402.98273.12865.1349
C53.70864.13984.33954.13991.40472.32672.23511.36732.19801.0778
O63.70024.29783.97944.29781.40471.49332.24772.27943.32492.1066
N72.47123.18942.53383.18932.32671.49331.34852.32193.35263.3257
C81.50932.15402.13192.15402.23512.24771.34851.44032.25923.3064
C92.64802.98263.50172.98271.36732.27942.32191.44031.07842.2528
H103.02963.12854.04713.12862.19803.32493.35262.25921.07842.7735
H114.76085.13485.41715.13491.07782.10663.32573.30642.25282.7735

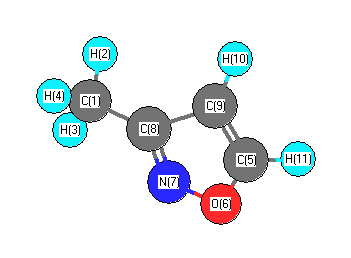
picture of 3-Methylisoxazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 N7 119.605 C1 C8 C9 127.706
H2 C1 H3 108.921 H2 C1 H4 108.830
H2 C1 C8 110.608 H3 C1 H4 108.919
H3 C1 C8 108.918 H4 C1 C8 110.608
C5 O6 N7 106.770 C5 C9 C8 105.484
C5 C9 H10 127.585 O6 C5 C9 110.619
O6 C5 H11 115.486 O6 N7 C8 104.438
N7 C8 C9 112.689 C8 C9 H10 126.931
C9 C5 H11 133.895
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability