return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SO2NH2 (methanesulfonamide)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-639.789605
Energy at 298.15K-639.797412
HF Energy-639.219380
Nuclear repulsion energy277.771521
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3523 3352 42.84      
2 A' 3194 3038 1.99      
3 A' 3108 2957 0.65      
4 A' 1692 1609 61.46      
5 A' 1565 1489 7.05      
6 A' 1478 1406 18.69      
7 A' 1208 1149 176.54      
8 A' 1067 1015 28.45      
9 A' 936 890 215.35      
10 A' 775 738 46.38      
11 A' 534 508 111.16      
12 A' 494 470 33.46      
13 A' 435 414 236.57      
14 A' 317 302 0.23      
15 A" 3655 3477 48.50      
16 A" 3206 3049 2.60      
17 A" 1566 1490 2.81      
18 A" 1455 1385 212.91      
19 A" 1103 1050 2.77      
20 A" 1041 990 0.06      
21 A" 433 412 0.27      
22 A" 345 328 3.08      
23 A" 259 246 0.00      
24 A" 160 153 18.65      

Unscaled Zero Point Vibrational Energy (zpe) 16774.4 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15957.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.15520 0.14656 0.14436

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.656 -0.046 0.000
S2 0.116 -0.137 0.000
N3 0.536 1.458 0.000
O4 0.536 -0.708 1.276
O5 0.536 -0.708 -1.276
H6 -2.059 -1.062 0.000
H7 -1.996 0.481 0.894
H8 -1.996 0.481 -0.894
H9 0.904 1.842 0.866
H10 0.904 1.842 -0.866

Atom - Atom Distances (Å)
  C1 S2 N3 O4 O5 H6 H7 H8 H9 H10
C11.77412.65772.62062.62061.09361.09191.09193.29593.2959
S21.77411.65001.45901.45902.36332.37562.37562.29942.2994
N32.65771.65002.51372.51373.61712.85732.85731.01601.0160
O42.62061.45902.51372.55152.91282.82303.53982.60813.3497
O52.62061.45902.51372.55152.91283.53982.82303.34972.6081
H61.09362.36333.61712.91282.91281.78451.78454.23784.2378
H71.09192.37562.85732.82303.53981.78451.78773.20363.6549
H81.09192.37562.85733.53982.82301.78451.78773.65493.2036
H93.29592.29941.01602.60813.34974.23783.20363.65491.7316
H103.29592.29941.01603.34972.60814.23783.65493.20361.7316

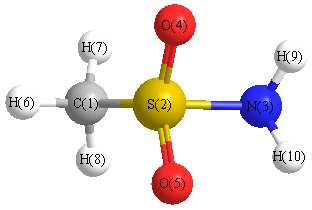
picture of methanesulfonamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 N3 101.761 C1 S2 O4 107.903
C1 S2 O5 107.903 S2 C1 H6 108.660
S2 C1 H7 109.660 S2 C1 H8 109.660
S2 N3 H9 117.196 S2 N3 H10 117.196
N3 S2 O4 107.747 N3 S2 O5 107.747
O4 S2 O5 121.952 H6 C1 H7 109.475
H6 C1 H8 109.475 H7 C1 H8 109.889
H9 N3 H10 116.902
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability