return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSSCH3 (dimethyl trisulfide)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-1266.350769
Energy at 298.15K-1266.357090
Nuclear repulsion energy366.401919
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3192 3037 4.23      
2 A 3176 3021 0.00      
3 A 3090 2940 2.76      
4 A 1581 1504 9.94      
5 A 1563 1486 9.72      
6 A 1452 1381 0.00      
7 A 1053 1002 0.47      
8 A 1049 998 5.60      
9 A 704 670 0.02      
10 A 502 478 0.44      
11 A 284 270 0.49      
12 A 195 185 1.61      
13 A 163 155 1.49      
14 A 66 63 2.44      
15 B 3192 3037 3.35      
16 B 3176 3021 3.82      
17 B 3090 2940 19.12      
18 B 1580 1503 12.39      
19 B 1562 1486 11.81      
20 B 1455 1384 2.21      
21 B 1052 1001 2.49      
22 B 1045 994 18.84      
23 B 703 669 1.54      
24 B 491 467 12.70      
25 B 288 274 0.94      
26 B 187 178 0.78      
27 B 102 97 4.29      

Unscaled Zero Point Vibrational Energy (zpe) 17996.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 17119.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14547 0.04717 0.04709

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.097
S2 0.000 1.635 -0.142
S3 0.000 -1.635 -0.142
C4 1.746 1.670 -0.684
C5 -1.746 -1.670 -0.684
H6 1.850 2.513 -1.374
H7 -1.850 -2.513 -1.374
H8 2.409 1.807 0.172
H9 1.987 0.740 -1.207
H10 -2.409 -1.807 0.172
H11 -1.987 -0.740 -1.207

Atom - Atom Distances (Å)
  S1 S2 S3 C4 C5 H6 H7 H8 H9 H10 H11
S12.05152.05153.00183.00183.98113.98113.15003.13113.15003.1311
S22.05153.27011.82853.77682.39044.70662.43502.42524.21263.2747
S32.05153.27013.77681.82854.70662.39044.21263.27472.43502.4252
C43.00181.82853.77684.83141.09475.55921.09171.09305.48444.4734
C53.00183.77681.82854.83145.55921.09475.48444.47341.09171.0930
H63.98112.39044.70661.09475.55926.24181.78941.78626.26035.0334
H73.98114.70662.39045.55921.09476.24186.26035.03341.78941.7862
H83.15002.43504.21261.09175.48441.78946.26031.79336.02205.2638
H93.13112.42523.27471.09304.47341.78625.03341.79335.26384.2403
H103.15004.21262.43505.48441.09176.26031.78946.02205.26381.7933
H113.13113.27472.42524.47341.09305.03341.78625.26384.24031.7933

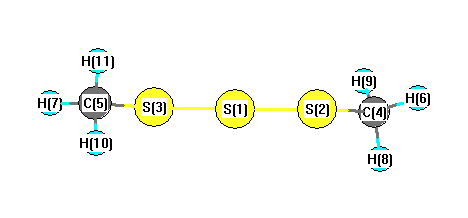
picture of dimethyl trisulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 101.214 S1 S3 C5 101.214
S2 S1 S3 105.692 S2 C4 H6 107.027
S2 C4 H8 110.443 S2 C4 H9 109.646
S3 C5 H7 107.027 S3 C5 H10 110.443
S3 C5 H11 109.646 H6 C4 H8 109.857
H6 C4 H9 109.466 H7 C5 H10 109.857
H7 C5 H11 109.466 H8 C4 H9 110.338
H10 C5 H11 110.338
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability