return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (orthodifluorobenzene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-426.807666
Energy at 298.15K-426.812487
Nuclear repulsion energy343.131103
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3244 3086 0.61      
2 A1 3225 3068 6.57      
3 A1 1610 1532 20.22      
4 A1 1557 1481 106.88      
5 A1 1337 1272 15.88      
6 A1 1320 1256 37.43      
7 A1 1233 1173 0.01      
8 A1 1046 995 5.08      
9 A1 770 733 17.67      
10 A1 599 570 3.73      
11 A1 274 261 0.56      
12 A2 941 895 0.00      
13 A2 874 831 0.00      
14 A2 710 676 0.00      
15 A2 575 547 0.00      
16 A2 194 184 0.00      
17 B1 946 900 16.14      
18 B1 783 745 104.61      
19 B1 486 462 2.75      
20 B1 308 293 0.03      
21 B2 3239 3081 1.36      
22 B2 3211 3054 1.14      
23 B2 1624 1545 3.73      
24 B2 1504 1431 9.36      
25 B2 1322 1257 3.72      
26 B2 1259 1198 17.65      
27 B2 1142 1086 14.55      
28 B2 891 848 12.10      
29 B2 549 523 2.58      
30 B2 440 418 0.70      

Unscaled Zero Point Vibrational Energy (zpe) 18605.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 17699.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.10644 0.07204 0.04296

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.701 -0.538
C2 0.000 -0.701 -0.538
C3 0.000 -1.398 0.673
C4 0.000 -0.702 1.891
C5 0.000 0.702 1.891
C6 0.000 1.398 0.673
F7 0.000 1.376 -1.736
F8 0.000 -1.376 -1.736
H9 0.000 -2.482 0.641
H10 0.000 -1.251 2.828
H11 0.000 1.251 2.828
H12 0.000 2.482 0.641

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C11.40222.42332.80542.42921.39701.37502.39793.39413.89103.41062.1356
C21.40221.39702.42922.80542.42332.39791.37502.13563.41063.89103.3941
C32.42331.39701.40342.42832.79643.67392.40861.08392.16053.41513.8799
C42.80542.42921.40341.40442.42834.18033.68922.17461.08562.16603.4204
C52.42922.80542.42831.40441.40343.68924.18033.42042.16601.08562.1746
C61.39702.42332.79642.42831.40342.40863.67393.87993.41512.16051.0839
F71.37502.39793.67394.18033.68922.40862.75224.53135.26594.56572.6216
F82.39791.37502.40863.68924.18033.67392.75222.62164.56575.26594.5313
H93.39412.13561.08392.17463.42043.87994.53132.62162.50944.32584.9632
H103.89103.41062.16051.08562.16603.41515.26594.56572.50942.50154.3258
H113.41063.89103.41512.16601.08562.16054.56575.26594.32582.50152.5094
H122.13563.39413.87993.42042.17461.08392.62164.53134.96324.32582.5094

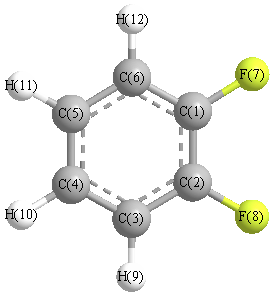
picture of orthodifluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.935 C1 C2 F8 119.402
C1 C6 C5 120.333 C1 C6 H12 118.276
C2 C1 C6 119.935 C2 C1 F7 119.402
C2 C3 C4 120.333 C2 C3 H9 118.276
C3 C2 F8 120.664 C3 C4 C5 119.732
C3 C4 H10 119.917 C4 C3 H9 121.391
C4 C5 C6 119.732 C4 C5 H11 120.350
C5 C4 H10 120.350 C5 C6 H12 121.391
C6 C1 F7 120.664 C6 C5 H11 119.917
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability