return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H6N2O2 ((E)-Azodioxymethane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-336.440483
Energy at 298.15K-336.448973
HF Energy-335.705474
Nuclear repulsion energy249.861613
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3244 3086 0.00      
2 Ag 3119 2967 0.00      
3 Ag 1595 1517 0.00      
4 Ag 1482 1410 0.00      
5 Ag 1437 1367 0.00      
6 Ag 1193 1135 0.00      
7 Ag 1027 977 0.00      
8 Ag 588 559 0.00      
9 Ag 530 504 0.00      
10 Ag 348 331 0.00      
11 Au 3197 3041 11.63      
12 Au 1573 1496 20.82      
13 Au 1171 1114 0.05      
14 Au 241 229 2.03      
15 Au 181 173 9.52      
16 Au 116 111 4.15      
17 Bg 3196 3041 0.00      
18 Bg 1572 1495 0.00      
19 Bg 1151 1095 0.00      
20 Bg 393 374 0.00      
21 Bg 146 139 0.00      
22 Bu 3244 3086 0.14      
23 Bu 3119 2967 7.60      
24 Bu 1679 1598 242.76      
25 Bu 1581 1504 9.73      
26 Bu 1484 1412 14.77      
27 Bu 1204 1145 22.44      
28 Bu 925 880 4.99      
29 Bu 485 461 51.91      
30 Bu 271 258 34.10      

Unscaled Zero Point Vibrational Energy (zpe) 20748.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 19737.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.15725 0.12522 0.07160

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.724 0.000
N2 0.000 -0.724 0.000
O3 -1.129 1.393 0.000
O4 1.129 -1.393 0.000
C5 1.389 1.275 0.000
C6 -1.389 -1.275 0.000
H7 1.268 2.356 0.000
H8 1.912 0.919 0.889
H9 1.912 0.919 -0.889
H10 -1.268 -2.356 0.000
H11 -1.912 -0.919 0.889
H12 -1.912 -0.919 -0.889

Atom - Atom Distances (Å)
  N1 N2 O3 O4 C5 C6 H7 H8 H9 H10 H11 H12
N11.44751.31272.39921.49462.43432.06692.11742.11743.33072.67262.6726
N21.44752.39921.31272.43431.49463.33072.67262.67262.06692.11742.1174
O31.31272.39923.58652.52112.68102.58343.20363.20363.75182.59752.5975
O42.39921.31273.58652.68102.52113.75182.59752.59752.58343.20363.2036
C51.49462.43432.52112.68103.77141.08761.09151.09154.49964.06194.0619
C62.43431.49462.68102.52113.77144.49964.06194.06191.08761.09151.0915
H72.06693.33072.58343.75181.08764.49961.80881.80885.35124.65024.6502
H82.11742.67263.20362.59751.09154.06191.80881.77894.65024.24184.5997
H92.11742.67263.20362.59751.09154.06191.80881.77894.65024.59974.2418
H103.33072.06693.75182.58344.49961.08765.35124.65024.65021.80881.8088
H112.67262.11742.59753.20364.06191.09154.65024.24184.59971.80881.7789
H122.67262.11742.59753.20364.06191.09154.65024.59974.24181.80881.7789

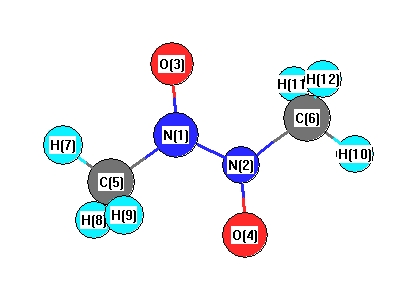
picture of (E)-Azodioxymethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 O4 120.653 N1 N2 C6 111.656
N1 C5 H7 105.255 N1 C5 H8 108.927
N1 C5 H9 108.927 N2 N1 O3 120.653
N2 N1 C5 111.656 N2 C6 H10 105.255
N2 C6 H11 108.927 N2 C6 H12 108.927
O3 N1 C5 127.692 O4 N2 C6 127.692
H7 C5 H8 112.214 H7 C5 H9 112.214
H8 C5 H9 109.155 H10 C6 H11 112.214
H10 C6 H12 112.214 H11 C6 H12 109.155
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability