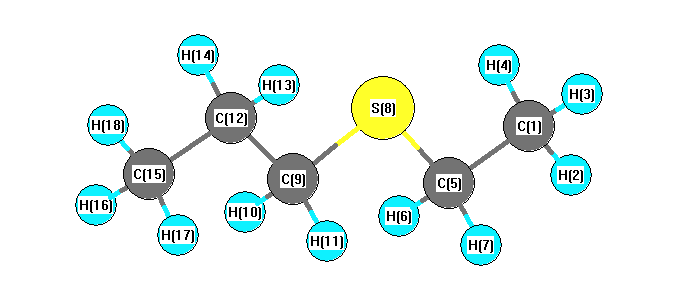
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3135 |
3007 |
18.60 |
|
|
|
| 2 |
A' |
3128 |
3001 |
28.10 |
|
|
|
| 3 |
A' |
3067 |
2943 |
21.09 |
|
|
|
| 4 |
A' |
3062 |
2937 |
18.86 |
|
|
|
| 5 |
A' |
3056 |
2932 |
22.21 |
|
|
|
| 6 |
A' |
3052 |
2928 |
26.04 |
|
|
|
| 7 |
A' |
3047 |
2923 |
11.65 |
|
|
|
| 8 |
A' |
1564 |
1500 |
4.85 |
|
|
|
| 9 |
A' |
1562 |
1499 |
9.27 |
|
|
|
| 10 |
A' |
1547 |
1485 |
2.99 |
|
|
|
| 11 |
A' |
1538 |
1475 |
2.82 |
|
|
|
| 12 |
A' |
1527 |
1465 |
11.63 |
|
|
|
| 13 |
A' |
1457 |
1397 |
0.13 |
|
|
|
| 14 |
A' |
1456 |
1397 |
19.52 |
|
|
|
| 15 |
A' |
1390 |
1334 |
0.58 |
|
|
|
| 16 |
A' |
1337 |
1283 |
14.42 |
|
|
|
| 17 |
A' |
1291 |
1239 |
15.26 |
|
|
|
| 18 |
A' |
1136 |
1090 |
5.38 |
|
|
|
| 19 |
A' |
1086 |
1042 |
18.05 |
|
|
|
| 20 |
A' |
1044 |
1001 |
0.19 |
|
|
|
| 21 |
A' |
993 |
953 |
3.15 |
|
|
|
| 22 |
A' |
912 |
875 |
2.53 |
|
|
|
| 23 |
A' |
772 |
741 |
1.03 |
|
|
|
| 24 |
A' |
691 |
663 |
0.32 |
|
|
|
| 25 |
A' |
391 |
375 |
0.53 |
|
|
|
| 26 |
A' |
304 |
292 |
0.78 |
|
|
|
| 27 |
A' |
278 |
266 |
0.79 |
|
|
|
| 28 |
A' |
105 |
101 |
0.35 |
|
|
|
| 29 |
A" |
3145 |
3017 |
20.66 |
|
|
|
| 30 |
A" |
3134 |
3007 |
48.77 |
|
|
|
| 31 |
A" |
3109 |
2983 |
0.90 |
|
|
|
| 32 |
A" |
3102 |
2976 |
6.97 |
|
|
|
| 33 |
A" |
3088 |
2963 |
3.30 |
|
|
|
| 34 |
A" |
1561 |
1498 |
10.82 |
|
|
|
| 35 |
A" |
1555 |
1492 |
11.93 |
|
|
|
| 36 |
A" |
1363 |
1308 |
0.11 |
|
|
|
| 37 |
A" |
1305 |
1252 |
0.17 |
|
|
|
| 38 |
A" |
1276 |
1225 |
0.04 |
|
|
|
| 39 |
A" |
1103 |
1058 |
1.73 |
|
|
|
| 40 |
A" |
1076 |
1033 |
0.58 |
|
|
|
| 41 |
A" |
900 |
863 |
0.08 |
|
|
|
| 42 |
A" |
819 |
786 |
9.97 |
|
|
|
| 43 |
A" |
770 |
738 |
6.42 |
|
|
|
| 44 |
A" |
246 |
236 |
0.06 |
|
|
|
| 45 |
A" |
240 |
231 |
0.03 |
|
|
|
| 46 |
A" |
118 |
113 |
0.91 |
|
|
|
| 47 |
A" |
67 |
64 |
0.37 |
|
|
|
| 48 |
A" |
41 |
39 |
0.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35972.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 34512.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.639 |
|
|
|
| 2 |
H |
0.221 |
|
|
|
| 3 |
H |
0.229 |
|
|
|
| 4 |
H |
0.229 |
|
|
|
| 5 |
C |
-0.603 |
|
|
|
| 6 |
H |
0.234 |
|
|
|
| 7 |
H |
0.234 |
|
|
|
| 8 |
S |
0.188 |
|
|
|
| 9 |
C |
-0.583 |
|
|
|
| 10 |
H |
0.230 |
|
|
|
| 11 |
H |
0.230 |
|
|
|
| 12 |
C |
-0.455 |
|
|
|
| 13 |
H |
0.231 |
|
|
|
| 14 |
H |
0.231 |
|
|
|
| 15 |
C |
-0.613 |
|
|
|
| 16 |
H |
0.211 |
|
|
|
| 17 |
H |
0.211 |
|
|
|
| 18 |
H |
0.216 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.978 |
-1.318 |
0.000 |
1.641 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.381 |
2.668 |
0.000 |
| y |
2.668 |
-46.699 |
0.000 |
| z |
0.000 |
0.000 |
-47.813 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.876 |
2.668 |
0.000 |
| y |
2.668 |
-1.102 |
0.000 |
| z |
0.000 |
0.000 |
-2.773 |
|
| Polar |
|---|
| 3z2-r2 | -5.546 |
|---|
| x2-y2 | 3.319 |
|---|
| xy | 2.668 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
355.202 |
| (<r2>)1/2 |
18.847 |