return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (Ethyl propyl sulfide)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-591.498636
Energy at 298.15K-591.511508
HF Energy-590.913842
Nuclear repulsion energy297.577379
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3164 3010 18.32      
2 A' 3157 3003 27.04      
3 A' 3098 2947 16.35      
4 A' 3091 2941 16.52      
5 A' 3090 2939 18.72      
6 A' 3082 2932 21.91      
7 A' 3077 2927 13.02      
8 A' 1606 1528 4.47      
9 A' 1605 1527 3.57      
10 A' 1593 1516 1.24      
11 A' 1587 1510 1.65      
12 A' 1579 1502 12.89      
13 A' 1496 1423 3.76      
14 A' 1494 1421 6.10      
15 A' 1428 1358 0.61      
16 A' 1378 1311 15.54      
17 A' 1337 1272 18.15      
18 A' 1153 1097 3.82      
19 A' 1109 1055 12.11      
20 A' 1031 981 0.34      
21 A' 984 936 2.07      
22 A' 908 864 1.58      
23 A' 781 743 1.27      
24 A' 700 666 0.61      
25 A' 400 380 0.48      
26 A' 309 294 0.71      
27 A' 285 271 0.73      
28 A' 110 105 0.33      
29 A" 3176 3021 20.53      
30 A" 3163 3009 47.15      
31 A" 3141 2988 0.45      
32 A" 3137 2984 8.04      
33 A" 3122 2970 4.51      
34 A" 1602 1524 7.47      
35 A" 1596 1518 7.89      
36 A" 1392 1324 0.16      
37 A" 1341 1276 0.01      
38 A" 1318 1254 0.00      
39 A" 1141 1086 1.12      
40 A" 1115 1061 0.20      
41 A" 927 882 0.07      
42 A" 839 798 6.13      
43 A" 785 747 4.25      
44 A" 257 245 0.02      
45 A" 251 239 0.10      
46 A" 125 119 0.94      
47 A" 78 74 0.41      
48 A" 48 46 0.34      

Unscaled Zero Point Vibrational Energy (zpe) 36591.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 34809.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.37743 0.03364 0.03182

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.385 2.298 0.000
H2 3.480 2.278 0.000
H3 2.044 2.837 0.889
H4 2.044 2.837 -0.889
C5 1.831 0.851 0.000
H6 2.178 0.315 -0.890
H7 2.178 0.315 0.890
S8 0.000 0.895 0.000
C9 -0.331 -0.904 0.000
H10 0.119 -1.358 -0.891
H11 0.119 -1.358 0.891
C12 -1.858 -1.164 0.000
H13 -2.302 -0.694 0.885
H14 -2.302 -0.694 -0.885
C15 -2.166 -2.683 0.000
H16 -1.738 -3.160 -0.889
H17 -1.738 -3.160 0.889
H18 -3.248 -2.858 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 S8 C9 H10 H11 C12 H13 H14 C15 H16 H17 H18
C11.09521.09411.09411.54962.18352.18352.76764.19944.39304.39305.47695.63105.63106.74786.89836.89837.6365
H21.09521.77911.77912.18062.51802.51803.74514.96525.03125.03126.35216.56136.56137.51657.58917.58918.4642
H31.09411.77911.77832.18633.08922.52552.95674.52004.94714.61575.65955.59995.87436.99957.30987.09027.8248
H41.09411.77911.77832.18632.52553.08922.95674.52004.61574.94715.65955.87435.59996.99957.09027.30987.8248
C51.54962.18062.18632.18631.09531.09531.83182.78522.93362.93364.20404.50054.50055.33595.44255.44256.2890
H62.18352.51803.08922.52551.09531.77992.42322.92832.65323.19554.39014.92354.59225.35325.23605.52996.3482
H72.18352.51802.52553.08921.09531.77992.42322.92833.19552.65324.39014.59224.92355.35325.52995.23606.3482
S82.76763.74512.95672.95671.83182.42322.42321.82922.42532.42532.77352.93382.93384.18264.50014.50014.9628
C94.19944.96524.52004.52002.78522.92832.92831.82921.09641.09641.54922.17062.17062.55592.80332.80333.5104
H104.39305.03124.94714.61572.93362.65323.19552.42531.09641.78202.17733.07502.51002.78792.58783.14093.7917
H114.39305.03124.61574.94712.93363.19552.65322.42531.09641.78202.17732.51003.07502.78793.14092.58783.7917
C125.47696.35215.65955.65954.20404.39014.39012.77351.54922.17732.17731.09611.09611.55002.18792.18792.1907
H135.63106.56135.59995.87434.50054.92354.59222.93382.17063.07502.51001.09611.77082.18133.08932.52892.5219
H145.63106.56135.87435.59994.50054.59224.92352.93382.17062.51003.07501.09611.77082.18132.52893.08932.5219
C156.74787.51656.99956.99955.33595.35325.35324.18262.55592.78792.78791.55002.18132.18131.09561.09561.0954
H166.89837.58917.30987.09025.44255.23605.52994.50012.80332.58783.14092.18793.08932.52891.09561.77801.7773
H176.89837.58917.09027.30985.44255.52995.23604.50012.80333.14092.58782.18792.52893.08931.09561.77801.7773
H187.63658.46427.82487.82486.28906.34826.34824.96283.51043.79173.79172.19072.52192.52191.09541.77731.7773

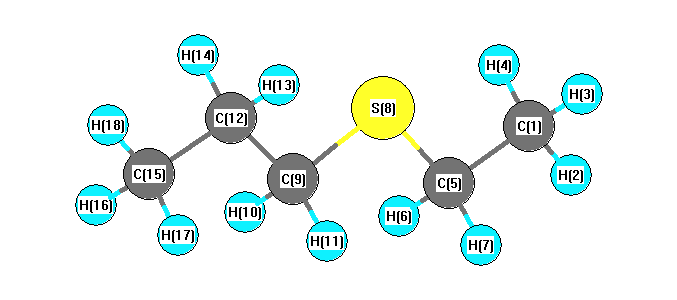
picture of Ethyl propyl sulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.108 C1 C5 H7 110.108
C1 C5 S8 109.585 H2 C1 H3 108.708
H2 C1 H4 108.708 H2 C1 C5 109.882
H3 C1 H4 108.715 H3 C1 C5 110.391
H4 C1 C5 110.391 C5 S8 C9 99.067
H6 C5 H7 108.689 H6 C5 S8 109.163
H7 C5 S8 109.163 S8 C9 H10 109.429
S8 C9 H11 109.429 S8 C9 C12 110.086
C9 C12 H13 109.085 C9 C12 H14 109.085
C9 C12 C15 111.118 H10 C9 H11 108.707
H10 C9 C12 109.584 H11 C9 C12 109.584
C12 C15 H16 110.406 C12 C15 H17 110.406
C12 C15 H18 110.638 H13 C12 H14 107.766
H13 C12 C15 109.859 H14 C12 C15 109.859
H16 C15 H17 108.473 H16 C15 H18 108.424
H17 C15 H18 108.424
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability