return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NNCH3 ((Z)-1,2-Dimethyldiazene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-187.391844
Energy at 298.15K-187.399141
HF Energy-186.969801
Nuclear repulsion energy118.643738
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3211 3055 2.21      
2 A1 3078 2928 7.40      
3 A1 1571 1494 0.03      
4 A1 1470 1398 14.12      
5 A1 1265 1203 6.01      
6 A1 1071 1019 12.74      
7 A1 757 720 1.10      
8 A1 359 341 0.64      
9 A2 3150 2996 0.00      
10 A2 1580 1503 0.00      
11 A2 1101 1048 0.00      
12 A2 440 419 0.00      
13 A2 101 97 0.00      
14 B1 3148 2995 30.43      
15 B1 1603 1525 25.19      
16 B1 969 922 0.98      
17 B1 202 192 0.08      
18 B2 3209 3053 24.40      
19 B2 3076 2926 0.12      
20 B2 1560 1484 13.19      
21 B2 1440 1370 0.68      
22 B2 1183 1125 4.41      
23 B2 827 787 14.80      
24 B2 576 548 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 18473.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 17573.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.50227 0.21424 0.15931

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.651 0.818
N2 0.000 -0.651 0.818
C3 0.000 1.389 -0.534
C4 0.000 -1.389 -0.534
H5 0.000 2.452 -0.295
H6 0.000 -2.452 -0.295
H7 0.893 1.133 -1.115
H8 -0.893 1.133 -1.115
H9 -0.893 -1.133 -1.115
H10 0.893 -1.133 -1.115

Atom - Atom Distances (Å)
  N1 N2 C3 C4 H5 H6 H7 H8 H9 H10
N11.30151.54062.44742.11743.29622.18362.18362.77812.7781
N21.30152.44741.54063.29622.11742.77812.77812.18362.1836
C31.54062.44742.77841.08893.84831.09591.09592.73812.7381
C42.44741.54062.77843.84831.08892.73812.73811.09591.0959
H52.11743.29621.08893.84834.90331.79121.79123.78423.7842
H63.29622.11743.84831.08894.90333.78423.78421.79121.7912
H72.18362.77811.09592.73811.79123.78421.78622.88552.2662
H82.18362.77811.09592.73811.79123.78421.78622.26622.8855
H92.77812.18362.73811.09593.78421.79122.88552.26621.7862
H102.77812.18362.73811.09593.78421.79122.26622.88551.7862

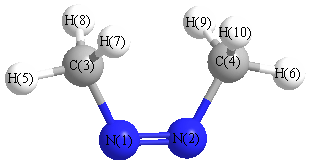
picture of (Z)-1,2-Dimethyldiazene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 118.641 N1 C3 H5 105.988
N1 C3 H7 110.698 N1 C3 H8 110.698
N2 N1 C3 118.641 N2 C4 H6 105.988
N2 C4 H9 110.698 N2 C4 H10 110.698
H5 C3 H7 110.134 H5 C3 H8 110.134
H6 C4 H9 110.134 H6 C4 H10 110.134
H7 C3 H8 109.160 H9 C4 H10 109.160
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability