return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-216.327851
Energy at 298.15K-216.335730
HF Energy-215.934891
Nuclear repulsion energy131.083145
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3183 3028 35.65      
2 A' 3180 3025 25.56      
3 A' 3102 2951 32.38      
4 A' 3100 2949 2.67      
5 A' 1612 1534 10.20      
6 A' 1594 1517 4.79      
7 A' 1503 1430 10.62      
8 A' 1444 1374 20.68      
9 A' 1262 1201 11.44      
10 A' 1214 1154 21.02      
11 A' 988 940 22.04      
12 A' 826 785 2.09      
13 A' 466 443 2.38      
14 A' 353 336 1.47      
15 A' 265 252 0.22      
16 A" 3179 3024 9.65      
17 A" 3174 3020 3.29      
18 A" 3097 2947 9.38      
19 A" 1591 1514 0.29      
20 A" 1583 1506 0.86      
21 A" 1487 1414 10.99      
22 A" 1455 1384 17.35      
23 A" 1185 1127 17.46      
24 A" 1004 955 1.53      
25 A" 971 923 0.11      
26 A" 376 358 6.90      
27 A" 231 220 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 21711.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20654.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.28007 0.26682 0.15622

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.294 0.253 0.000
F2 -0.907 1.048 0.000
H3 1.149 0.937 0.000
C4 0.294 -0.592 1.278
C5 0.294 -0.592 -1.278
H6 1.205 -1.194 1.334
H7 1.205 -1.194 -1.334
H8 0.237 0.057 2.157
H9 0.237 0.057 -2.157
H10 -0.577 -1.255 1.274
H11 -0.577 -1.255 -1.274

Atom - Atom Distances (Å)
  C1 F2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.44061.09521.53261.53262.16892.16892.16622.16622.15752.1575
F21.44062.05982.40182.40183.35723.35722.63482.63482.65282.6528
H31.09522.05982.16892.16892.51472.51472.50152.50153.06703.0670
C41.53262.40182.16892.55671.09362.83121.09363.49621.09412.7769
C51.53262.40182.16892.55672.83121.09363.49621.09362.77691.0941
H62.16893.35722.51471.09362.83122.66731.78353.83221.78383.1588
H72.16893.35722.51472.83121.09362.66733.83221.78353.15881.7838
H82.16622.63482.50151.09363.49621.78353.83224.31311.77823.7618
H92.16622.63482.50153.49621.09363.83221.78354.31313.76181.7782
H102.15752.65283.06701.09412.77691.78383.15881.77823.76182.5479
H112.15752.65283.06702.77691.09413.15881.78383.76181.77822.5479

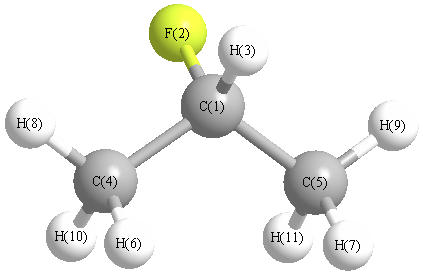
picture of 2-Fluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H8 110.020 C1 C4 H10 109.313
C1 C5 H7 110.237 C1 C5 H9 110.020
C1 C5 H11 109.313 F2 C1 H3 107.865
F2 C1 C4 107.729 F2 C1 C5 107.729
H3 C1 C4 110.139 H3 C1 C5 110.139
C4 C1 C5 113.051 H7 C5 H9 109.257
H7 C5 H11 109.247 H8 C4 H10 108.741
H9 C5 H11 108.741
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability