Jump to
S1C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -186.235090 |
| Energy at 298.15K | -186.238237 |
| HF Energy | -185.848834 |
| Nuclear repulsion energy | 101.479883 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3687 |
3495 |
25.00 |
|
|
|
| 2 |
A |
3577 |
3391 |
0.00 |
|
|
|
| 3 |
A |
2410 |
2285 |
0.00 |
|
|
|
| 4 |
A |
1750 |
1659 |
0.00 |
|
|
|
| 5 |
A |
1207 |
1144 |
4.74 |
|
|
|
| 6 |
A |
826 |
783 |
0.00 |
|
|
|
| 7 |
A |
544 |
515 |
12.43 |
|
|
|
| 8 |
A |
514 |
487 |
0.00 |
|
|
|
| 9 |
A |
261 |
248 |
18.86 |
|
|
|
| 10 |
A |
39 |
37 |
415.22 |
|
|
|
| 11 |
B |
3687 |
3495 |
25.00 |
|
|
|
| 12 |
B |
3582 |
3396 |
39.91 |
|
|
|
| 13 |
B |
1755 |
1664 |
48.68 |
|
|
|
| 14 |
B |
1413 |
1340 |
127.98 |
|
|
|
| 15 |
B |
1207 |
1144 |
4.74 |
|
|
|
| 16 |
B |
544 |
515 |
12.43 |
|
|
|
| 17 |
B |
261 |
248 |
18.86 |
|
|
|
| 18 |
B |
39 |
37 |
415.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13650.9 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 12941.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.007 |
0.610 |
-0.006 |
| C2 |
-0.007 |
-0.610 |
-0.006 |
| N3 |
-0.007 |
1.983 |
-0.039 |
| N4 |
0.007 |
-1.983 |
-0.039 |
| H5 |
-0.398 |
2.474 |
0.763 |
| H6 |
0.793 |
2.455 |
-0.458 |
| H7 |
0.398 |
-2.474 |
0.763 |
| H8 |
-0.793 |
-2.455 |
-0.458 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2210 | 1.3730 | 2.5937 | 2.0567 | 2.0552 | 3.2034 | 3.1998 |
C2 | 1.2210 | | 2.5937 | 1.3730 | 3.2034 | 3.1998 | 2.0567 | 2.0552 | N3 | 1.3730 | 2.5937 | | 3.9661 | 1.0186 | 1.0189 | 4.5470 | 4.5263 | N4 | 2.5937 | 1.3730 | 3.9661 | | 4.5470 | 4.5263 | 1.0186 | 1.0189 | H5 | 2.0567 | 3.2034 | 1.0186 | 4.5470 | | 1.7063 | 5.0124 | 5.0935 | H6 | 2.0552 | 3.1998 | 1.0189 | 4.5263 | 1.7063 | | 5.0935 | 5.1591 | H7 | 3.2034 | 2.0567 | 4.5470 | 1.0186 | 5.0124 | 5.0935 | | 1.7063 | H8 | 3.1998 | 2.0552 | 4.5263 | 1.0189 | 5.0935 | 5.1591 | 1.7063 | |
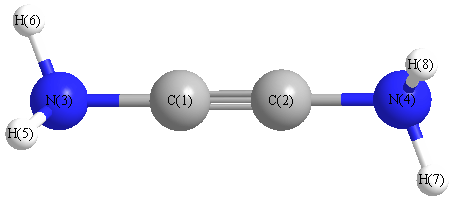 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
178.198 |
|
C1 |
N3 |
H5 |
117.858 |
| C1 |
N3 |
H6 |
117.697 |
|
C2 |
C1 |
N3 |
178.198 |
| C2 |
N4 |
H7 |
117.858 |
|
C2 |
N4 |
H8 |
117.697 |
| H5 |
N3 |
H6 |
113.736 |
|
H7 |
N4 |
H8 |
113.736 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -186.235090 |
| Energy at 298.15K | -186.238210 |
| HF Energy | -185.848828 |
| Nuclear repulsion energy | 101.478701 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3577 |
3391 |
0.00 |
|
|
|
| 2 |
A1 |
2410 |
2285 |
0.00 |
|
|
|
| 3 |
A1 |
1750 |
1659 |
0.00 |
|
|
|
| 4 |
A1 |
826 |
783 |
0.00 |
|
|
|
| 5 |
B1 |
514 |
487 |
0.00 |
|
|
|
| 6 |
B2 |
3582 |
3396 |
39.93 |
|
|
|
| 7 |
B2 |
1755 |
1664 |
48.67 |
|
|
|
| 8 |
B2 |
1413 |
1340 |
127.99 |
|
|
|
| 9 |
E |
3687 |
3496 |
25.03 |
|
|
|
| 9 |
E |
3687 |
3496 |
25.03 |
|
|
|
| 10 |
E |
1207 |
1144 |
4.74 |
|
|
|
| 10 |
E |
1207 |
1144 |
4.74 |
|
|
|
| 11 |
E |
544 |
516 |
12.45 |
|
|
|
| 11 |
E |
544 |
516 |
12.45 |
|
|
|
| 12 |
E |
261 |
248 |
18.67 |
|
|
|
| 12 |
E |
261 |
248 |
18.67 |
|
|
|
| 13 |
E |
32 |
30 |
415.42 |
|
|
|
| 13 |
E |
32 |
30 |
415.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13644.3 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 12934.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.611 |
| C2 |
0.000 |
0.000 |
-0.611 |
| N3 |
0.000 |
0.000 |
1.961 |
| N4 |
0.000 |
0.000 |
-1.961 |
| H5 |
0.000 |
0.864 |
2.485 |
| H6 |
0.000 |
-0.864 |
2.485 |
| H7 |
0.864 |
0.000 |
-2.485 |
| H8 |
-0.864 |
0.000 |
-2.485 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2226 | 1.3498 | 2.5724 | 2.0630 | 2.0630 | 3.2143 | 3.2143 |
C2 | 1.2226 | | 2.5724 | 1.3498 | 3.2143 | 3.2143 | 2.0630 | 2.0630 | N3 | 1.3498 | 2.5724 | | 3.9222 | 1.0101 | 1.0101 | 4.5290 | 4.5290 | N4 | 2.5724 | 1.3498 | 3.9222 | | 4.5290 | 4.5290 | 1.0101 | 1.0101 | H5 | 2.0630 | 3.2143 | 1.0101 | 4.5290 | | 1.7274 | 5.1176 | 5.1176 | H6 | 2.0630 | 3.2143 | 1.0101 | 4.5290 | 1.7274 | | 5.1176 | 5.1176 | H7 | 3.2143 | 2.0630 | 4.5290 | 1.0101 | 5.1176 | 5.1176 | | 1.7274 | H8 | 3.2143 | 2.0630 | 4.5290 | 1.0101 | 5.1176 | 5.1176 | 1.7274 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.232 |
| C1 |
N3 |
H6 |
121.232 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.232 |
|
C2 |
N4 |
H8 |
121.232 |
| H5 |
N3 |
H6 |
117.536 |
|
H7 |
N4 |
H8 |
117.536 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
