Jump to
S1C2
Energy calculated at B97D3/3-21G*
| | hartrees |
|---|
| Energy at 0K | -186.890522 |
| Energy at 298.15K | -186.894913 |
| HF Energy | -186.890522 |
| Nuclear repulsion energy | 101.869475 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3497 |
3497 |
5.80 |
|
|
|
| 2 |
A |
3414 |
3414 |
0.00 |
|
|
|
| 3 |
A |
2332 |
2332 |
0.00 |
|
|
|
| 4 |
A |
1674 |
1674 |
0.00 |
|
|
|
| 5 |
A |
1169 |
1169 |
3.48 |
|
|
|
| 6 |
A |
845 |
845 |
0.00 |
|
|
|
| 7 |
A |
708 |
708 |
40.11 |
|
|
|
| 8 |
A |
592 |
592 |
0.00 |
|
|
|
| 9 |
A |
377 |
377 |
323.00 |
|
|
|
| 10 |
A |
270 |
270 |
39.48 |
|
|
|
| 11 |
B |
3497 |
3497 |
5.80 |
|
|
|
| 12 |
B |
3425 |
3425 |
0.11 |
|
|
|
| 13 |
B |
1684 |
1684 |
20.43 |
|
|
|
| 14 |
B |
1449 |
1449 |
100.12 |
|
|
|
| 15 |
B |
1169 |
1169 |
3.48 |
|
|
|
| 16 |
B |
708 |
708 |
40.10 |
|
|
|
| 17 |
B |
377 |
377 |
322.94 |
|
|
|
| 18 |
B |
270 |
270 |
39.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13727.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13727.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/3-21G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.003 |
0.611 |
-0.009 |
| C2 |
-0.003 |
-0.611 |
-0.009 |
| N3 |
-0.003 |
1.970 |
-0.035 |
| N4 |
0.003 |
-1.970 |
-0.035 |
| H5 |
-0.405 |
2.471 |
0.763 |
| H6 |
0.795 |
2.449 |
-0.464 |
| H7 |
0.405 |
-2.471 |
0.763 |
| H8 |
-0.795 |
-2.449 |
-0.464 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2214 | 1.3595 | 2.5808 | 2.0545 | 2.0527 | 3.2018 | 3.1946 |
C2 | 1.2214 | | 2.5808 | 1.3595 | 3.2018 | 3.1946 | 2.0545 | 2.0527 | N3 | 1.3595 | 2.5808 | | 3.9399 | 1.0242 | 1.0245 | 4.5301 | 4.5099 | N4 | 2.5808 | 1.3595 | 3.9399 | | 4.5301 | 4.5099 | 1.0242 | 1.0245 | H5 | 2.0545 | 3.2018 | 1.0242 | 4.5301 | | 1.7159 | 5.0070 | 5.0854 | H6 | 2.0527 | 3.1946 | 1.0245 | 4.5099 | 1.7159 | | 5.0854 | 5.1496 | H7 | 3.2018 | 2.0545 | 4.5301 | 1.0242 | 5.0070 | 5.0854 | | 1.7159 | H8 | 3.1946 | 2.0527 | 4.5099 | 1.0245 | 5.0854 | 5.1496 | 1.7159 | |
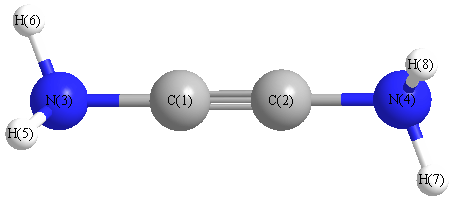 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
178.783 |
|
C1 |
N3 |
H5 |
118.387 |
| C1 |
N3 |
H6 |
118.184 |
|
C2 |
C1 |
N3 |
178.783 |
| C2 |
N4 |
H7 |
118.387 |
|
C2 |
N4 |
H8 |
118.184 |
| H5 |
N3 |
H6 |
113.764 |
|
H7 |
N4 |
H8 |
113.764 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.378 |
|
|
|
| 2 |
C |
0.378 |
|
|
|
| 3 |
N |
-0.968 |
|
|
|
| 4 |
N |
-0.968 |
|
|
|
| 5 |
H |
0.295 |
|
|
|
| 6 |
H |
0.295 |
|
|
|
| 7 |
H |
0.295 |
|
|
|
| 8 |
H |
0.295 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.001 |
0.001 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.296 |
-0.006 |
0.000 |
| y |
-0.006 |
-9.602 |
0.000 |
| z |
0.000 |
0.000 |
-23.297 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.847 |
-0.006 |
0.000 |
| y |
-0.006 |
13.694 |
0.000 |
| z |
0.000 |
0.000 |
-6.848 |
|
| Polar |
|---|
| 3z2-r2 | -13.695 |
|---|
| x2-y2 | -13.694 |
|---|
| xy | -0.006 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.958 |
0.000 |
0.000 |
| y |
0.000 |
9.178 |
0.000 |
| z |
0.000 |
0.000 |
1.958 |
<r2> (average value of r
2) Å
2
| <r2> |
96.911 |
| (<r2>)1/2 |
9.844 |
Jump to
S1C1
Energy calculated at B97D3/3-21G*
| | hartrees |
|---|
| Energy at 0K | -186.890512 |
| Energy at 298.15K | -186.894894 |
| HF Energy | -186.890512 |
| Nuclear repulsion energy | 101.866780 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3415 |
3415 |
0.00 |
|
|
|
| 2 |
A1 |
2332 |
2332 |
0.00 |
|
|
|
| 3 |
A1 |
1674 |
1674 |
0.00 |
|
|
|
| 4 |
A1 |
845 |
845 |
0.00 |
|
|
|
| 5 |
B1 |
590 |
590 |
0.00 |
|
|
|
| 6 |
B2 |
3426 |
3426 |
0.10 |
|
|
|
| 7 |
B2 |
1684 |
1684 |
20.44 |
|
|
|
| 8 |
B2 |
1449 |
1449 |
100.24 |
|
|
|
| 9 |
E |
3498 |
3498 |
5.82 |
|
|
|
| 9 |
E |
3498 |
3498 |
5.82 |
|
|
|
| 10 |
E |
1168 |
1168 |
3.46 |
|
|
|
| 10 |
E |
1168 |
1168 |
3.46 |
|
|
|
| 11 |
E |
708 |
708 |
40.06 |
|
|
|
| 11 |
E |
708 |
708 |
40.06 |
|
|
|
| 12 |
E |
375 |
375 |
322.21 |
|
|
|
| 12 |
E |
375 |
375 |
322.21 |
|
|
|
| 13 |
E |
269 |
269 |
40.32 |
|
|
|
| 13 |
E |
269 |
269 |
40.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13724.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13724.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/3-21G*
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.613 |
| C2 |
0.000 |
0.000 |
-0.613 |
| N3 |
0.000 |
0.000 |
1.946 |
| N4 |
0.000 |
0.000 |
-1.946 |
| H5 |
0.000 |
0.866 |
2.483 |
| H6 |
0.000 |
-0.866 |
2.483 |
| H7 |
0.866 |
0.000 |
-2.483 |
| H8 |
-0.866 |
0.000 |
-2.483 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2270 | 1.3328 | 2.5598 | 2.0607 | 2.0607 | 3.2157 | 3.2157 |
C2 | 1.2270 | | 2.5598 | 1.3328 | 3.2157 | 3.2157 | 2.0607 | 2.0607 | N3 | 1.3328 | 2.5598 | | 3.8925 | 1.0189 | 1.0189 | 4.5136 | 4.5136 | N4 | 2.5598 | 1.3328 | 3.8925 | | 4.5136 | 4.5136 | 1.0189 | 1.0189 | H5 | 2.0607 | 3.2157 | 1.0189 | 4.5136 | | 1.7316 | 5.1157 | 5.1157 | H6 | 2.0607 | 3.2157 | 1.0189 | 4.5136 | 1.7316 | | 5.1157 | 5.1157 | H7 | 3.2157 | 2.0607 | 4.5136 | 1.0189 | 5.1157 | 5.1157 | | 1.7316 | H8 | 3.2157 | 2.0607 | 4.5136 | 1.0189 | 5.1157 | 5.1157 | 1.7316 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.818 |
| C1 |
N3 |
H6 |
121.818 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.818 |
|
C2 |
N4 |
H8 |
121.818 |
| H5 |
N3 |
H6 |
116.363 |
|
H7 |
N4 |
H8 |
116.363 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.378 |
|
|
|
| 2 |
C |
0.378 |
|
|
|
| 3 |
N |
-0.968 |
|
|
|
| 4 |
N |
-0.968 |
|
|
|
| 5 |
H |
0.295 |
|
|
|
| 6 |
H |
0.295 |
|
|
|
| 7 |
H |
0.295 |
|
|
|
| 8 |
H |
0.295 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.296 |
0.000 |
0.000 |
| y |
0.000 |
-23.296 |
0.000 |
| z |
0.000 |
0.000 |
-9.604 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.846 |
0.000 |
0.000 |
| y |
0.000 |
-6.846 |
0.000 |
| z |
0.000 |
0.000 |
13.692 |
|
| Polar |
|---|
| 3z2-r2 | 27.384 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.958 |
0.000 |
0.000 |
| y |
0.000 |
1.958 |
0.000 |
| z |
0.000 |
0.000 |
9.177 |
<r2> (average value of r
2) Å
2
| <r2> |
96.917 |
| (<r2>)1/2 |
9.845 |
