return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H7N (Cyclobutanecarbonitrile)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-247.016649
Energy at 298.15K-247.024666
HF Energy-246.445404
Nuclear repulsion energy208.183676
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3209 3053 27.44      
2 A' 3189 3034 5.67      
3 A' 3134 2981 3.65      
4 A' 3129 2977 12.10      
5 A' 3127 2975 7.30      
6 A' 2139 2035 1.52      
7 A' 1592 1514 1.00      
8 A' 1578 1501 5.23      
9 A' 1396 1328 0.71      
10 A' 1324 1259 3.55      
11 A' 1271 1210 0.22      
12 A' 1164 1107 1.81      
13 A' 1067 1015 0.06      
14 A' 946 900 0.30      
15 A' 915 871 2.47      
16 A' 752 715 1.71      
17 A' 600 571 0.75      
18 A' 540 513 1.47      
19 A' 296 282 0.65      
20 A' 143 136 2.29      
21 A" 3199 3043 11.66      
22 A" 3128 2976 26.32      
23 A" 1569 1492 2.67      
24 A" 1331 1266 3.33      
25 A" 1308 1245 0.48      
26 A" 1296 1233 0.35      
27 A" 1236 1176 1.36      
28 A" 1061 1010 0.29      
29 A" 940 894 1.15      
30 A" 901 857 1.46      
31 A" 819 779 1.27      
32 A" 563 535 0.13      
33 A" 198 189 2.76      

Unscaled Zero Point Vibrational Energy (zpe) 24528.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23334.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.31850 0.07735 0.06830

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.666 2.129 0.000
C2 0.749 1.371 0.000
C3 -0.350 0.415 0.000
C4 -0.911 -1.712 0.000
C5 -0.350 -0.729 1.101
C6 -0.350 -0.729 -1.101
H7 -1.318 0.925 0.000
H8 -2.005 -1.710 0.000
H9 -0.532 -2.734 0.000
H10 0.679 -0.988 1.366
H11 -0.948 -0.553 1.996
H12 0.679 -0.988 -1.366
H13 -0.948 -0.553 -1.996

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13
N11.18922.64594.62533.66613.66613.21795.31135.33703.54364.24403.54364.2440
C21.18921.45703.50192.61332.61332.11524.13264.30102.72743.25112.72743.2511
C32.64591.45702.19951.58741.58741.09482.69323.15452.21202.29792.21202.2979
C44.62533.50192.19951.57971.57972.66821.09331.09062.21772.30842.21772.3084
C53.66612.61331.58741.57972.20272.21042.21702.29571.09331.09072.68583.1598
C63.66612.61331.58741.57972.20272.21042.21702.29572.68583.15981.09331.0907
H73.21792.11521.09482.66822.21042.21042.72303.74303.08482.51163.08482.5116
H85.31134.13262.69321.09332.21702.21702.72301.79383.09652.53773.09652.5377
H95.33704.30103.15451.09062.29572.29573.74301.79382.52632.98592.52632.9859
H103.54362.72742.21202.21771.09332.68583.08483.09652.52631.79802.73233.7605
H114.24403.25112.29792.30841.09073.15982.51162.53772.98591.79803.76053.9926
H123.54362.72742.21202.21772.68581.09333.08483.09652.52632.73233.76051.7980
H134.24403.25112.29792.30843.15981.09072.51162.53772.98593.76053.99261.7980

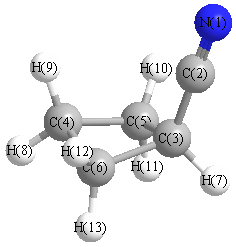
picture of Cyclobutanecarbonitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 178.543 C2 C3 C5 118.213
C2 C3 C6 118.213 C2 C3 H7 111.183
C3 C5 C4 87.971 C3 C5 H10 109.829
C3 C5 H11 116.973 C3 C6 C4 87.971
C3 C6 H12 109.829 C3 C6 H13 116.973
C4 C5 H10 110.818 C4 C5 H11 118.492
C4 C6 H12 110.818 C4 C6 H13 118.492
C5 C3 C6 87.861 C5 C3 H7 109.618
C5 C4 C6 88.405 C5 C4 H8 110.761
C5 C4 H9 117.393 C6 C3 H7 109.618
C6 C4 H8 110.761 C6 C4 H9 117.393
H8 C4 H9 110.439 H10 C5 H11 110.820
H12 C6 H13 110.820
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability