return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H12N2 (1,2-Butanediamine)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-266.439398
Energy at 298.15K-266.454394
HF Energy-265.855512
Nuclear repulsion energy261.717172
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3537 3365 2.55      
2 A 3528 3356 0.37      
3 A 3425 3258 10.33      
4 A 3419 3253 2.47      
5 A 3188 3032 15.40      
6 A 3150 2997 31.45      
7 A 3143 2990 24.12      
8 A 3110 2959 39.38      
9 A 3094 2943 8.11      
10 A 3077 2927 19.60      
11 A 2991 2845 83.59      
12 A 2963 2819 75.12      
13 A 1747 1662 27.56      
14 A 1723 1639 42.17      
15 A 1614 1536 1.81      
16 A 1610 1531 7.91      
17 A 1602 1524 3.61      
18 A 1580 1503 4.84      
19 A 1492 1420 6.67      
20 A 1475 1403 7.96      
21 A 1453 1382 6.54      
22 A 1436 1366 4.61      
23 A 1398 1330 0.78      
24 A 1362 1295 1.76      
25 A 1323 1259 2.88      
26 A 1306 1242 2.66      
27 A 1229 1170 8.12      
28 A 1175 1117 11.47      
29 A 1129 1074 0.34      
30 A 1105 1051 12.25      
31 A 1063 1011 7.86      
32 A 1033 983 1.31      
33 A 986 938 9.09      
34 A 921 876 8.22      
35 A 911 866 50.56      
36 A 844 803 148.27      
37 A 818 778 71.78      
38 A 780 742 22.50      
39 A 623 593 22.81      
40 A 451 429 6.37      
41 A 401 381 20.62      
42 A 380 362 69.68      
43 A 322 306 20.93      
44 A 269 256 6.52      
45 A 245 233 22.41      
46 A 228 217 4.21      
47 A 171 162 10.63      
48 A 83 79 1.51      

Unscaled Zero Point Vibrational Energy (zpe) 37454.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35630.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.18246 0.07352 0.06400

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.900 -0.609 -0.695
H2 1.715 0.209 -1.284
H3 2.891 -0.863 -0.718
C4 -2.421 -0.309 -0.122
H5 -3.049 -1.116 -0.516
H6 -2.779 -0.052 0.882
H7 -2.558 0.562 -0.773
N8 0.081 1.504 -0.308
H9 0.526 2.280 0.193
H10 -0.847 1.799 -0.620
C11 -0.937 -0.757 -0.073
H12 -0.863 -1.678 0.519
H13 -0.552 -0.975 -1.071
C14 1.443 -0.265 0.680
H15 2.062 0.494 1.196
H16 1.436 -1.168 1.301
C17 0.002 0.287 0.570
H18 -0.368 0.511 1.587

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 H7 N8 H9 H10 C11 H12 H13 C14 H15 H16 C17 H18
N11.02451.02284.36924.97844.96904.60942.81443.32043.65372.90833.20192.50761.48972.19502.12462.45033.4067
H21.02451.68844.32745.00444.99584.31762.30202.80863.08773.07143.66872.56642.03892.52022.94242.52523.5599
H31.02281.68845.37355.94875.94645.63163.69604.03734.58953.88304.03523.46262.09942.48732.50703.36474.2208
C44.36924.32745.37351.09551.09661.09583.09523.93522.67751.55102.17142.19933.94644.74174.20012.58962.7941
H54.97845.00445.94871.09551.77681.76734.08684.98143.65452.18772.48362.56214.72565.62584.83963.52943.7755
H64.96904.99585.94641.09661.77681.77893.46664.10323.06882.19122.53913.10244.23184.88234.38032.81922.5742
H74.60944.31765.63161.09581.76731.77892.83953.65912.11612.20363.09212.54434.33545.02224.82102.90333.2191
N82.81442.30203.69603.09524.08683.46662.83951.02581.02272.49043.42052.66932.44102.68473.40061.50252.1861
H93.32042.80864.03733.93524.98144.10323.65911.02581.66663.38164.20763.65452.74872.56083.73452.09502.4233
H103.65373.08774.58952.67753.65453.06882.11611.02271.66662.61553.65932.82563.34573.66974.20812.10332.6002
C112.90833.07143.88301.55102.18772.19122.20362.49043.38162.61551.09771.09172.54423.48882.77291.54442.1647
H123.20193.66874.03522.17142.48362.53913.09213.42054.20763.65931.09771.76652.70913.70592.48122.14772.4852
H132.50762.56643.46262.19932.56213.10242.54432.66933.65452.82561.09171.76652.74783.75923.10132.14273.0504
C141.48972.03892.09943.94644.72564.23184.33542.44102.74873.34572.54422.70912.74781.10711.09621.54642.1685
H152.19502.52022.48734.74175.62584.88235.02222.68472.56083.66973.48883.70593.75921.10711.77922.16322.4618
H162.12462.94242.50704.20014.83964.38034.82103.40063.73454.20812.77292.48123.10131.09621.77922.16972.4808
C172.45032.52523.36472.58963.52942.81922.90331.50252.09502.10331.54442.14772.14271.54642.16322.16971.1053
H183.40673.55994.22082.79413.77552.57423.21912.18612.42332.60022.16472.48523.05042.16852.46182.48081.1053

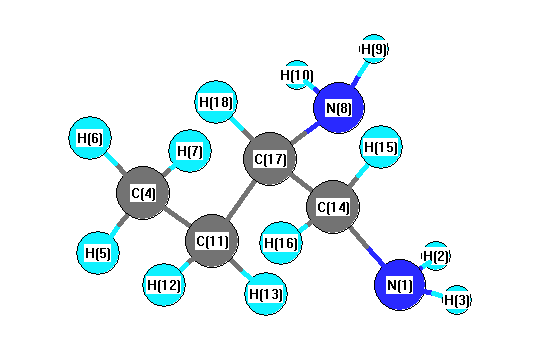
picture of 1,2-Butanediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C14 H15 114.605 N1 C14 H16 109.554
N1 C14 C17 107.601 H2 N1 H3 111.115
H2 N1 C14 106.917 H3 N1 C14 112.013
C4 C11 H12 108.924 C4 C11 H13 111.471
C4 C11 C17 113.568 H5 C4 H6 108.302
H5 C4 H7 107.512 H5 C4 C11 110.326
H6 C4 H7 108.472 H6 C4 C11 110.542
H7 C4 C11 111.572 N8 C17 C11 109.640
N8 C17 C14 106.367 N8 C17 H18 113.037
H9 N8 H10 108.894 H9 N8 C17 110.499
H10 N8 C17 111.388 C11 C17 C14 110.800
C11 C17 H18 108.425 H12 C11 H13 107.578
H12 C11 C17 107.556 H13 C11 C17 107.509
C14 C17 H18 108.574 H15 C14 H16 107.715
H15 C14 C17 108.074 H16 C14 C17 109.192
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability