Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -216.320285 |
| Energy at 298.15K | -216.328192 |
| HF Energy | -215.928203 |
| Nuclear repulsion energy | 126.490643 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3166 |
3012 |
22.45 |
|
|
|
| 2 |
A' |
3111 |
2960 |
22.11 |
|
|
|
| 3 |
A' |
3088 |
2938 |
30.73 |
|
|
|
| 4 |
A' |
3084 |
2934 |
9.82 |
|
|
|
| 5 |
A' |
1630 |
1551 |
0.76 |
|
|
|
| 6 |
A' |
1607 |
1528 |
5.76 |
|
|
|
| 7 |
A' |
1596 |
1518 |
2.13 |
|
|
|
| 8 |
A' |
1505 |
1431 |
16.44 |
|
|
|
| 9 |
A' |
1486 |
1413 |
4.65 |
|
|
|
| 10 |
A' |
1397 |
1329 |
0.32 |
|
|
|
| 11 |
A' |
1171 |
1114 |
0.73 |
|
|
|
| 12 |
A' |
1082 |
1029 |
44.82 |
|
|
|
| 13 |
A' |
1028 |
978 |
2.47 |
|
|
|
| 14 |
A' |
914 |
869 |
11.76 |
|
|
|
| 15 |
A' |
443 |
421 |
4.71 |
|
|
|
| 16 |
A' |
261 |
248 |
4.67 |
|
|
|
| 17 |
A" |
3169 |
3015 |
61.41 |
|
|
|
| 18 |
A" |
3145 |
2992 |
0.00 |
|
|
|
| 19 |
A" |
3128 |
2975 |
22.79 |
|
|
|
| 20 |
A" |
1606 |
1528 |
8.58 |
|
|
|
| 21 |
A" |
1390 |
1323 |
0.71 |
|
|
|
| 22 |
A" |
1331 |
1266 |
0.01 |
|
|
|
| 23 |
A" |
1243 |
1182 |
0.75 |
|
|
|
| 24 |
A" |
956 |
910 |
2.29 |
|
|
|
| 25 |
A" |
805 |
766 |
2.46 |
|
|
|
| 26 |
A" |
247 |
235 |
0.00 |
|
|
|
| 27 |
A" |
144 |
137 |
3.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21866.2 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20801.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.112 |
-0.792 |
0.000 |
| C2 |
0.000 |
0.735 |
0.000 |
| C3 |
-1.481 |
1.184 |
0.000 |
| F4 |
1.496 |
-1.164 |
0.000 |
| H5 |
-0.369 |
-1.209 |
0.890 |
| H6 |
-0.369 |
-1.209 |
-0.890 |
| H7 |
0.517 |
1.123 |
-0.884 |
| H8 |
0.517 |
1.123 |
0.884 |
| H9 |
-1.553 |
2.276 |
0.000 |
| H10 |
-1.998 |
0.805 |
-0.889 |
| H11 |
-1.998 |
0.805 |
0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5314 | 2.5382 | 1.4328 | 1.0951 | 1.0951 | 2.1471 | 2.1471 | 3.4904 | 2.7921 | 2.7921 |
C2 | 1.5314 | | 1.5472 | 2.4181 | 2.1705 | 2.1705 | 1.0949 | 1.0949 | 2.1873 | 2.1880 | 2.1880 | C3 | 2.5382 | 1.5472 | | 3.7916 | 2.7849 | 2.7849 | 2.1852 | 2.1852 | 1.0940 | 1.0958 | 1.0958 | F4 | 1.4328 | 2.4181 | 3.7916 | | 2.0675 | 2.0675 | 2.6404 | 2.6404 | 4.5968 | 4.1085 | 4.1085 | H5 | 1.0951 | 2.1705 | 2.7849 | 2.0675 | | 1.7809 | 3.0614 | 2.4944 | 3.7866 | 3.1429 | 2.5909 | H6 | 1.0951 | 2.1705 | 2.7849 | 2.0675 | 1.7809 | | 2.4944 | 3.0614 | 3.7866 | 2.5909 | 3.1429 | H7 | 2.1471 | 1.0949 | 2.1852 | 2.6404 | 3.0614 | 2.4944 | | 1.7687 | 2.5288 | 2.5346 | 3.0932 | H8 | 2.1471 | 1.0949 | 2.1852 | 2.6404 | 2.4944 | 3.0614 | 1.7687 | | 2.5288 | 3.0932 | 2.5346 | H9 | 3.4904 | 2.1873 | 1.0940 | 4.5968 | 3.7866 | 3.7866 | 2.5288 | 2.5288 | | 1.7746 | 1.7746 | H10 | 2.7921 | 2.1880 | 1.0958 | 4.1085 | 3.1429 | 2.5909 | 2.5346 | 3.0932 | 1.7746 | | 1.7773 | H11 | 2.7921 | 2.1880 | 1.0958 | 4.1085 | 2.5909 | 3.1429 | 3.0932 | 2.5346 | 1.7746 | 1.7773 | |
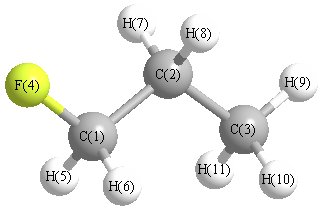 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.066 |
|
C1 |
C2 |
H7 |
108.541 |
| C1 |
C2 |
H8 |
108.541 |
|
C2 |
C1 |
F4 |
109.282 |
| C2 |
C1 |
H5 |
110.355 |
|
C2 |
C1 |
H6 |
110.355 |
| C2 |
C3 |
H9 |
110.640 |
|
C2 |
C3 |
H10 |
110.597 |
| C2 |
C3 |
H11 |
110.597 |
|
C3 |
C2 |
H7 |
110.426 |
| C3 |
C2 |
H8 |
110.426 |
|
F4 |
C1 |
H5 |
109.008 |
| F4 |
C1 |
H6 |
109.008 |
|
H5 |
C1 |
H6 |
108.804 |
| H7 |
C2 |
H8 |
107.747 |
|
H9 |
C3 |
H10 |
108.270 |
| H9 |
C3 |
H11 |
108.270 |
|
H10 |
C3 |
H11 |
108.383 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -216.321398 |
| Energy at 298.15K | |
| HF Energy | -215.928978 |
| Nuclear repulsion energy | 129.646604 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3192 |
3036 |
14.66 |
|
|
|
| 2 |
A |
3166 |
3012 |
43.72 |
|
|
|
| 3 |
A |
3149 |
2996 |
19.64 |
|
|
|
| 4 |
A |
3138 |
2985 |
30.65 |
|
|
|
| 5 |
A |
3103 |
2952 |
10.74 |
|
|
|
| 6 |
A |
3093 |
2943 |
13.05 |
|
|
|
| 7 |
A |
3092 |
2941 |
37.29 |
|
|
|
| 8 |
A |
1625 |
1546 |
0.94 |
|
|
|
| 9 |
A |
1617 |
1538 |
6.42 |
|
|
|
| 10 |
A |
1599 |
1521 |
7.20 |
|
|
|
| 11 |
A |
1581 |
1504 |
2.78 |
|
|
|
| 12 |
A |
1498 |
1425 |
9.58 |
|
|
|
| 13 |
A |
1477 |
1405 |
10.00 |
|
|
|
| 14 |
A |
1446 |
1376 |
0.42 |
|
|
|
| 15 |
A |
1366 |
1300 |
0.53 |
|
|
|
| 16 |
A |
1318 |
1254 |
0.38 |
|
|
|
| 17 |
A |
1228 |
1168 |
1.19 |
|
|
|
| 18 |
A |
1163 |
1107 |
7.74 |
|
|
|
| 19 |
A |
1087 |
1034 |
22.54 |
|
|
|
| 20 |
A |
1015 |
965 |
21.15 |
|
|
|
| 21 |
A |
970 |
922 |
7.16 |
|
|
|
| 22 |
A |
876 |
834 |
3.62 |
|
|
|
| 23 |
A |
806 |
767 |
1.29 |
|
|
|
| 24 |
A |
490 |
466 |
2.49 |
|
|
|
| 25 |
A |
318 |
303 |
2.13 |
|
|
|
| 26 |
A |
252 |
239 |
3.60 |
|
|
|
| 27 |
A |
160 |
152 |
3.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21911.3 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20844.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.806 |
0.556 |
0.283 |
| C2 |
-0.614 |
0.700 |
-0.276 |
| C3 |
-1.477 |
-0.520 |
0.124 |
| F4 |
1.346 |
-0.697 |
-0.167 |
| H5 |
1.456 |
1.365 |
-0.063 |
| H6 |
0.786 |
0.553 |
1.378 |
| H7 |
-0.556 |
0.766 |
-1.368 |
| H8 |
-1.050 |
1.633 |
0.098 |
| H9 |
-2.458 |
-0.490 |
-0.360 |
| H10 |
-0.950 |
-1.427 |
-0.180 |
| H11 |
-1.623 |
-0.547 |
1.210 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5330 | 2.5288 | 1.4370 | 1.0944 | 1.0948 | 2.1502 | 2.1535 | 3.4867 | 2.6882 | 2.8237 |
C2 | 1.5330 | | 1.5473 | 2.4098 | 2.1854 | 2.1719 | 1.0954 | 1.0956 | 2.1958 | 2.1556 | 2.1870 | C3 | 2.5288 | 1.5473 | | 2.8436 | 3.4922 | 2.8005 | 2.1753 | 2.1954 | 1.0940 | 1.0924 | 1.0958 | F4 | 1.4370 | 2.4098 | 2.8436 | | 2.0677 | 2.0647 | 2.6833 | 3.3527 | 3.8141 | 2.4086 | 3.2756 | H5 | 1.0944 | 2.1854 | 3.4922 | 2.0677 | | 1.7846 | 2.4719 | 2.5260 | 4.3417 | 3.6874 | 3.8418 | H6 | 1.0948 | 2.1719 | 2.8005 | 2.0647 | 1.7846 | | 3.0633 | 2.4846 | 3.8244 | 3.0588 | 2.6530 | H7 | 2.1502 | 1.0954 | 2.1753 | 2.6833 | 2.4719 | 3.0633 | | 1.7737 | 2.4926 | 2.5255 | 3.0842 | H8 | 2.1535 | 1.0956 | 2.1954 | 3.3527 | 2.5260 | 2.4846 | 1.7737 | | 2.5882 | 3.0743 | 2.5140 | H9 | 3.4867 | 2.1958 | 1.0940 | 3.8141 | 4.3417 | 3.8244 | 2.4926 | 2.5882 | | 1.7848 | 1.7793 | H10 | 2.6882 | 2.1556 | 1.0924 | 2.4086 | 3.6874 | 3.0588 | 2.5255 | 3.0743 | 1.7848 | | 1.7769 | H11 | 2.8237 | 2.1870 | 1.0958 | 3.2756 | 3.8418 | 2.6530 | 3.0842 | 2.5140 | 1.7793 | 1.7769 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
110.358 |
|
C1 |
C2 |
H7 |
108.637 |
| C1 |
C2 |
H8 |
108.888 |
|
C2 |
C1 |
F4 |
108.423 |
| C2 |
C1 |
H5 |
111.471 |
|
C2 |
C1 |
H6 |
110.371 |
| C2 |
C3 |
H9 |
111.314 |
|
C2 |
C3 |
H10 |
108.256 |
| C2 |
C3 |
H11 |
110.505 |
|
C3 |
C2 |
H7 |
109.608 |
| C3 |
C2 |
H8 |
111.185 |
|
F4 |
C1 |
H5 |
108.783 |
| F4 |
C1 |
H6 |
108.521 |
|
H5 |
C1 |
H6 |
109.208 |
| H7 |
C2 |
H8 |
108.098 |
|
H9 |
C3 |
H10 |
109.438 |
| H9 |
C3 |
H11 |
108.689 |
|
H10 |
C3 |
H11 |
108.593 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
