return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5F (Fluorobenzene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-328.376638
Energy at 298.15K-328.382238
HF Energy-327.739368
Nuclear repulsion energy266.543387
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3240 3082 0.02      
2 A1 3222 3065 12.56      
3 A1 3198 3043 0.00      
4 A1 1622 1543 51.25      
5 A1 1544 1469 43.54      
6 A1 1297 1234 43.49      
7 A1 1230 1170 3.81      
8 A1 1055 1004 0.44      
9 A1 1024 974 2.27      
10 A1 830 790 12.83      
11 A1 538 512 2.67      
12 A2 949 903 0.00      
13 A2 847 806 0.00      
14 A2 433 412 0.00      
15 B1 936 890 0.00      
16 B1 903 859 15.26      
17 B1 768 730 104.78      
18 B1 684 650 9.28      
19 B1 515 490 7.69      
20 B1 246 234 0.14      
21 B2 3236 3079 3.75      
22 B2 3209 3053 7.22      
23 B2 1602 1524 8.86      
24 B2 1502 1429 2.23      
25 B2 1377 1310 0.24      
26 B2 1323 1258 0.06      
27 B2 1237 1177 0.87      
28 B2 1101 1048 6.25      
29 B2 653 621 0.07      
30 B2 391 372 2.33      

Unscaled Zero Point Vibrational Energy (zpe) 20355.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 19364.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.18772 0.08298 0.05754

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
F1 0.000 0.000 2.324
C2 0.000 0.000 0.940
C3 0.000 1.218 0.255
C4 0.000 -1.218 0.255
C5 0.000 1.217 -1.150
C6 0.000 -1.217 -1.150
C7 0.000 0.000 -1.856
H8 0.000 2.145 0.819
H9 0.000 -2.145 0.819
H10 0.000 2.160 -1.687
H11 0.000 -2.160 -1.687
H12 0.000 0.000 -2.942

Atom - Atom Distances (Å)
  F1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
F11.38412.40082.40083.68053.68054.18032.62032.62034.55614.55615.2664
C21.38411.39731.39732.41792.41792.79622.14812.14813.40143.40143.8822
C32.40081.39732.43611.40482.81082.43761.08463.40972.15903.89703.4217
C42.40081.39732.43612.81081.40482.43763.40971.08463.89702.15903.4217
C53.68052.41791.40482.81082.43301.40692.17623.89521.08623.41932.1665
C63.68052.41792.81081.40482.43301.40693.89522.17623.41931.08622.1665
C74.18032.79622.43762.43761.40691.40693.42863.42862.16692.16691.0860
H82.62032.14811.08463.40972.17623.89523.42864.28942.50624.98134.3296
H92.62032.14813.40971.08463.89522.17623.42864.28944.98132.50624.3296
H104.55613.40142.15903.89701.08623.41932.16692.50624.98134.32052.4983
H114.55613.40143.89702.15903.41931.08622.16694.98132.50624.32052.4983
H125.26643.88223.42173.42172.16652.16651.08604.32964.32962.49832.4983

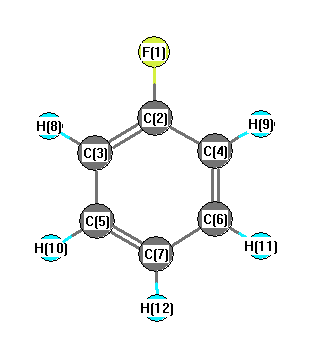
picture of Fluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
F1 C2 C3 119.342 F1 C2 C4 119.342
C2 C3 C5 119.279 C2 C3 H8 119.353
C2 C4 C6 119.279 C2 C4 H9 119.353
C3 C2 C4 121.316 C3 C5 C7 120.215
C3 C5 H10 119.612 C4 C6 C7 120.215
C4 C6 H11 119.612 C5 C3 H8 121.367
C5 C7 C6 119.696 C5 C7 H12 120.152
C6 C4 H9 121.367 C6 C7 H12 120.152
C7 C5 H10 120.173 C7 C6 H11 120.173
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability