Jump to
S1C2
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -355.208277 |
| Energy at 298.15K | -355.213584 |
| HF Energy | -354.536558 |
| Nuclear repulsion energy | 228.668312 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3707 |
3514 |
67.64 |
|
|
|
| 2 |
A' |
3566 |
3380 |
71.20 |
|
|
|
| 3 |
A' |
3504 |
3322 |
38.63 |
|
|
|
| 4 |
A' |
1746 |
1655 |
130.60 |
|
|
|
| 5 |
A' |
1706 |
1617 |
109.15 |
|
|
|
| 6 |
A' |
1665 |
1578 |
153.80 |
|
|
|
| 7 |
A' |
1426 |
1352 |
8.75 |
|
|
|
| 8 |
A' |
1332 |
1262 |
77.05 |
|
|
|
| 9 |
A' |
1126 |
1068 |
195.54 |
|
|
|
| 10 |
A' |
1100 |
1043 |
98.09 |
|
|
|
| 11 |
A' |
741 |
702 |
3.75 |
|
|
|
| 12 |
A' |
608 |
576 |
61.67 |
|
|
|
| 13 |
A' |
537 |
509 |
3.62 |
|
|
|
| 14 |
A' |
425 |
403 |
9.27 |
|
|
|
| 15 |
A' |
269 |
255 |
11.06 |
|
|
|
| 16 |
A" |
839 |
795 |
0.08 |
|
|
|
| 17 |
A" |
668 |
634 |
273.81 |
|
|
|
| 18 |
A" |
641 |
607 |
52.91 |
|
|
|
| 19 |
A" |
579 |
549 |
188.49 |
|
|
|
| 20 |
A" |
431 |
409 |
50.99 |
|
|
|
| 21 |
A" |
115 |
109 |
4.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13365.0 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 12670.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.744 |
0.000 |
| C2 |
-0.047 |
-0.794 |
0.000 |
| O3 |
-1.102 |
-1.461 |
0.000 |
| O4 |
1.047 |
1.415 |
0.000 |
| O5 |
-1.266 |
1.277 |
0.000 |
| N6 |
1.233 |
-1.275 |
0.000 |
| H7 |
1.399 |
-2.273 |
0.000 |
| H8 |
2.005 |
-0.621 |
0.000 |
| H9 |
-1.186 |
2.270 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5385 | 2.4648 | 1.2434 | 1.3736 | 2.3661 | 3.3256 | 2.4260 | 1.9327 |
C2 | 1.5385 | | 1.2478 | 2.4645 | 2.4026 | 1.3676 | 2.0685 | 2.0597 | 3.2685 | O3 | 2.4648 | 1.2478 | | 3.5896 | 2.7426 | 2.3417 | 2.6287 | 3.2184 | 3.7319 | O4 | 1.2434 | 2.4645 | 3.5896 | | 2.3172 | 2.6965 | 3.7045 | 2.2501 | 2.3914 | O5 | 1.3736 | 2.4026 | 2.7426 | 2.3172 | | 3.5719 | 4.4387 | 3.7821 | 0.9964 | N6 | 2.3661 | 1.3676 | 2.3417 | 2.6965 | 3.5719 | | 1.0113 | 1.0124 | 4.2921 | H7 | 3.3256 | 2.0685 | 2.6287 | 3.7045 | 4.4387 | 1.0113 | | 1.7599 | 5.2269 | H8 | 2.4260 | 2.0597 | 3.2184 | 2.2501 | 3.7821 | 1.0124 | 1.7599 | | 4.3063 | H9 | 1.9327 | 3.2685 | 3.7319 | 2.3914 | 0.9964 | 4.2921 | 5.2269 | 4.3063 | |
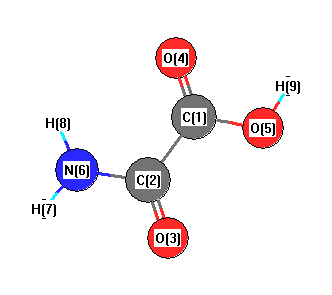 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
124.081 |
|
C1 |
C2 |
N6 |
108.871 |
| C1 |
O5 |
H9 |
108.221 |
|
C2 |
C1 |
O4 |
124.393 |
| C2 |
C1 |
O5 |
111.063 |
|
C2 |
N6 |
H7 |
120.057 |
| C2 |
N6 |
H8 |
119.109 |
|
O3 |
C2 |
N6 |
127.048 |
| O4 |
C1 |
O5 |
124.544 |
|
H7 |
N6 |
H8 |
120.834 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -355.213798 |
| Energy at 298.15K | -355.219321 |
| Nuclear repulsion energy | 230.090320 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3689 |
3497 |
76.63 |
|
|
|
| 2 |
A' |
3549 |
3365 |
86.54 |
|
|
|
| 3 |
A' |
3371 |
3195 |
95.18 |
|
|
|
| 4 |
A' |
1761 |
1669 |
79.16 |
|
|
|
| 5 |
A' |
1746 |
1655 |
199.14 |
|
|
|
| 6 |
A' |
1666 |
1579 |
36.53 |
|
|
|
| 7 |
A' |
1407 |
1334 |
91.84 |
|
|
|
| 8 |
A' |
1337 |
1268 |
456.10 |
|
|
|
| 9 |
A' |
1176 |
1115 |
30.10 |
|
|
|
| 10 |
A' |
1116 |
1058 |
1.18 |
|
|
|
| 11 |
A' |
755 |
716 |
13.90 |
|
|
|
| 12 |
A' |
631 |
598 |
7.89 |
|
|
|
| 13 |
A' |
558 |
529 |
1.20 |
|
|
|
| 14 |
A' |
418 |
396 |
5.67 |
|
|
|
| 15 |
A' |
279 |
264 |
43.75 |
|
|
|
| 16 |
A" |
826 |
783 |
11.93 |
|
|
|
| 17 |
A" |
762 |
722 |
164.81 |
|
|
|
| 18 |
A" |
672 |
637 |
33.29 |
|
|
|
| 19 |
A" |
636 |
603 |
327.10 |
|
|
|
| 20 |
A" |
448 |
425 |
2.77 |
|
|
|
| 21 |
A" |
158 |
150 |
5.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13478.9 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 12778.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.024 |
-0.790 |
0.000 |
| C2 |
0.000 |
0.754 |
0.000 |
| O3 |
-1.108 |
1.356 |
0.000 |
| O4 |
1.059 |
-1.470 |
0.000 |
| O5 |
-1.251 |
-1.289 |
0.000 |
| N6 |
1.243 |
1.284 |
0.000 |
| H7 |
1.385 |
2.286 |
0.000 |
| H8 |
2.032 |
0.646 |
0.000 |
| H9 |
-1.859 |
-0.488 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5445 | 2.4260 | 1.2386 | 1.3686 | 2.4058 | 3.3640 | 2.4691 | 1.9061 |
C2 | 1.5445 | | 1.2608 | 2.4632 | 2.3955 | 1.3509 | 2.0653 | 2.0348 | 2.2356 | O3 | 2.4260 | 1.2608 | | 3.5608 | 2.6484 | 2.3518 | 2.6611 | 3.2191 | 1.9911 | O4 | 1.2386 | 2.4632 | 3.5608 | | 2.3171 | 2.7597 | 3.7697 | 2.3288 | 3.0781 | O5 | 1.3686 | 2.3955 | 2.6484 | 2.3171 | | 3.5829 | 4.4416 | 3.8107 | 1.0047 | N6 | 2.4058 | 1.3509 | 2.3518 | 2.7597 | 3.5829 | | 1.0121 | 1.0147 | 3.5720 | H7 | 3.3640 | 2.0653 | 2.6611 | 3.7697 | 4.4416 | 1.0121 | | 1.7626 | 4.2684 | H8 | 2.4691 | 2.0348 | 3.2191 | 2.3288 | 3.8107 | 1.0147 | 1.7626 | | 4.0525 | H9 | 1.9061 | 2.2356 | 1.9911 | 3.0781 | 1.0047 | 3.5720 | 4.2684 | 4.0525 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.378 |
|
C1 |
C2 |
N6 |
112.211 |
| C1 |
O5 |
H9 |
105.839 |
|
C2 |
C1 |
O4 |
124.151 |
| C2 |
C1 |
O5 |
110.489 |
|
C2 |
N6 |
H7 |
121.186 |
| C2 |
N6 |
H8 |
117.974 |
|
O3 |
C2 |
N6 |
128.411 |
| O4 |
C1 |
O5 |
125.360 |
|
H7 |
N6 |
H8 |
120.841 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
