return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (Thiophene, tetrahydro-3-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-590.338863
Energy at 298.15K-590.350883
HF Energy-589.758507
Nuclear repulsion energy311.398418
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3176 3022 13.02      
2 A 3165 3011 21.41      
3 A 3163 3009 10.80      
4 A 3150 2997 24.74      
5 A 3144 2991 6.88      
6 A 3120 2968 12.37      
7 A 3111 2959 33.52      
8 A 3098 2947 13.35      
9 A 3088 2938 6.86      
10 A 3080 2930 15.52      
11 A 1609 1531 6.53      
12 A 1597 1519 2.38      
13 A 1588 1511 5.69      
14 A 1579 1502 2.87      
15 A 1569 1493 5.69      
16 A 1488 1416 10.52      
17 A 1433 1363 3.18      
18 A 1394 1326 6.46      
19 A 1370 1304 0.54      
20 A 1360 1294 10.73      
21 A 1307 1244 6.47      
22 A 1282 1219 4.26      
23 A 1223 1164 2.56      
24 A 1195 1137 0.51      
25 A 1134 1079 1.91      
26 A 1122 1067 1.24      
27 A 1078 1025 0.73      
28 A 999 951 1.03      
29 A 986 938 0.38      
30 A 945 899 2.48      
31 A 930 885 1.91      
32 A 875 833 0.64      
33 A 801 762 2.41      
34 A 731 696 4.97      
35 A 694 660 2.93      
36 A 621 590 1.15      
37 A 514 489 0.14      
38 A 390 371 0.13      
39 A 365 347 0.09      
40 A 250 238 0.11      
41 A 201 191 0.18      
42 A 120 114 2.84      

Unscaled Zero Point Vibrational Energy (zpe) 32022.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30462.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.13864 0.08639 0.07091

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 -1.465 -0.443 -0.264
C2 1.850 -0.339 -0.853
H3 1.123 -0.278 -1.670
H4 2.613 0.436 -0.994
H5 2.338 -1.319 -0.899
C6 -0.037 -1.160 0.665
H7 0.207 -2.139 0.244
H8 -0.307 -1.272 1.719
C9 1.139 -0.158 0.508
H10 1.858 -0.314 1.323
C11 -0.740 1.257 -0.320
H12 -1.492 1.982 0.002
H13 -0.433 1.486 -1.343
C14 0.487 1.244 0.634
H15 0.147 1.392 1.666
H16 1.192 2.041 0.373

Atom - Atom Distances (Å)
  S1 C2 H3 H4 H5 C6 H7 H8 C9 H10 C11 H12 H13 C14 H15 H16
S13.36842.94984.23453.95371.84802.43592.44212.73143.68471.84882.43982.43922.73173.11333.6930
C23.36841.09481.09611.09602.55752.67293.48431.54602.17593.08864.15742.96372.56383.49872.7568
H32.94981.09481.78441.77572.75242.82273.81042.18103.08142.76593.83922.37512.83323.85593.0911
H44.23451.09611.78441.77893.51003.73504.33622.18592.54933.51714.49753.24072.79663.75072.5422
H53.95371.09601.77571.77892.84852.55333.72192.18302.48544.05545.13573.96813.51344.32783.7712
C61.84802.55752.75243.51002.84851.09411.09391.55362.17712.70243.52483.34512.46012.74723.4413
H72.43592.67292.82273.73502.55331.09411.78662.20592.68683.57094.46454.00943.41703.80754.2970
H82.44213.48433.81044.33623.72191.09391.78662.19122.40033.27763.86574.12362.85282.70343.8780
C92.73141.54602.18102.18592.18301.55362.20592.19121.09752.49383.42872.93291.55082.17422.2035
H103.68472.17593.08142.54932.48542.17712.68682.40031.09753.45194.27033.94932.18642.44042.6253
C111.84883.08862.76593.51714.05542.70243.57093.27762.49383.45191.09271.09271.55432.17922.1976
H122.43984.15743.83924.49755.13573.52484.46453.86573.42874.27031.09271.78202.20442.40872.7101
H132.43922.96372.37513.24073.96813.34514.00944.12362.93293.94931.09271.78202.19423.06602.4280
C142.73172.56382.83322.79663.51342.46013.41702.85281.55082.18641.55432.20442.19421.09661.0961
H153.11333.49873.85593.75074.32782.74723.80752.70342.17422.44042.17922.40873.06601.09661.7848
H163.69302.75683.09112.54223.77123.44134.29703.87802.20352.62532.19762.71012.42801.09611.7848

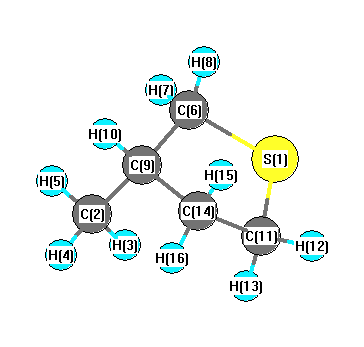
picture of Thiophene, tetrahydro-3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C6 H7 109.070 S1 C6 H8 109.538
S1 C6 C9 106.511 S1 C11 H12 109.378
S1 C11 H13 109.339 S1 C11 C14 106.458
C2 C9 C6 111.196 C2 C9 H10 109.628
C2 C9 C14 111.762 H3 C2 H4 109.061
H3 C2 H5 108.292 H3 C2 C9 110.184
H4 C2 H5 108.484 H4 C2 C9 110.495
H5 C2 C9 110.270 C6 S1 C11 93.946
C6 C9 H10 109.205 C6 C9 C14 104.829
H7 C6 H8 109.475 H7 C6 C9 111.669
H8 C6 C9 110.513 C9 C14 C11 106.855
C9 C14 H15 109.217 C9 C14 H16 111.553
H10 C9 C14 110.114 C11 C14 H15 109.363
C11 C14 H16 110.837 H12 C11 H13 109.255
H12 C11 C14 111.580 H13 C11 C14 110.770
H15 C14 H16 108.971
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability